


For family planning
Первый шаг в планировании беременности и семьи.
Тест ENVISION carrier screening даёт самую надёжную и подробную информацию о здоровье вашей будущей семьи
Что такое ENVISION
carrier screening test?
Скрининг Envision показывает, являетесь ли вы и ваш партнёр носителями изменённых генов, которые можно передать ребёнку.
Если ребёнок унаследует изменённый ген от обоих родителей, развиваются серьёзные генетические нарушения, такие как муковисцидоз, спинальная мышечная атрофия и тяжёлые гематологические заболевания.
Большинство людей являются носителями скрытых изменений в структуре генов и не знают об этом.
Эти изменения не влияют на ваше здоровье, но у вашего ребёнка повышен риск развития заболевания.
Из-за характера наследования этих нарушений невозможно предсказать, у каких родителей и в какой беременности родится больной ребёнок.
Предотвратить заболевания с аутосомно-рецессивным и X-сцепленным рецессивным наследованием можно только путём определения статуса носительства, то есть с помощью скрининга носительства.
РИСКИ, КОТОРЫХ МЫ НЕ ВИДИМ
 детей с генетическими нарушениями рождаются у родителей без семейной истории заболевания и без его симптомов.
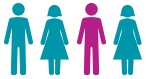
1 из 4 человек — носитель гена как минимум одного заболевания.*
* Клиническое исследование Lazarin et al. (2011)
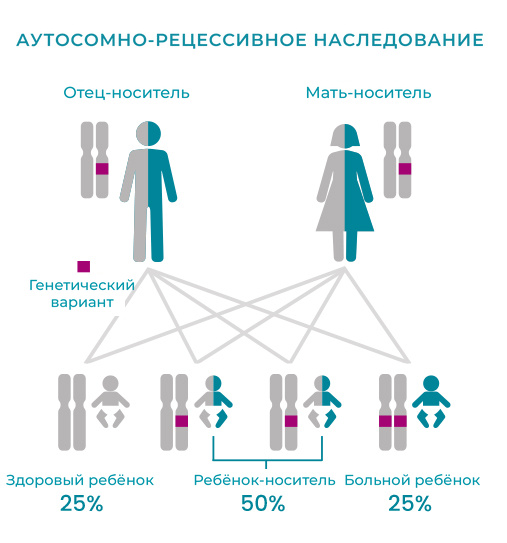
Аутосомно-рецессивное наследование означает, что для развития болезни человек должен унаследовать изменённый ген от обоих родителей.
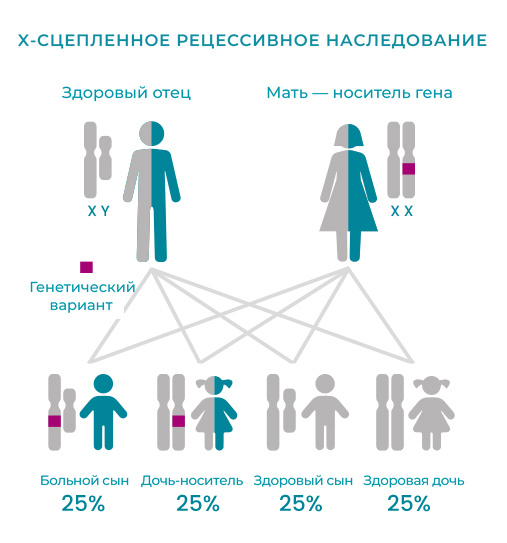
X-сцепленное рецессивное наследование означает, что ген находится на X-хромосоме.
У женщин-носительниц обычно нет симптомов, так как на второй X-хромосоме у них есть функциональная копия гена.
Поскольку у мужчин одна X-хромосома, унаследованный мутантный ген означает, что они будут больны
Здоровье малыша — в ваших генах.
Планируя будущее с партнёром, прекрасно иметь как можно больше общего.
Однако когда речь идёт о генах, различия желательны и полезны.

Кому подходит тест
ENVISION carrier screening?

Парам, планирующим семью

Парам на раннем сроке беременности
Если вы ещё не проходили тест Envision, сделайте его сейчас: полученная информация относится к этой и ко всем последующим беременностям.

Парам, проходящим ЭКО и/или имевшим неудачную попытку ЭКО
Проходя этот одновременно сложный и прекрасный путь, важно сделать всё, чтобы свести риск неудачи к минимуму.

Людям, в семье которых есть генетические заболевания
Если в семье есть случай генетического заболевания, необходимо проверить, есть ли у вашего ребёнка риск его унаследовать и насколько он высок.

Определённым этническим группам с повышенным риском генетических заболеваний
В некоторых группах населения доказана более высокая частота генетических заболеваний; их представителям рекомендуется тест Envision как обязательная часть планирования семьи.

Каждому, кто хочет узнать, является ли он носителем изменённых генов, которые могут привести к заболеванию у потомства
Скрининг носительства нужен всем беременным на раннем сроке и всем женщинам, планирующим беременность.
Obstet Gynecol. 2017.
Что анализирует тест ENVISION carrier screening
и какую технологию он использует?
Как мы выбирали
заболевания для анализа?
Все заболевания, рекомендованные Американской коллегией медицинской генетики (ACMG) и Американской коллегией акушеров и гинекологов (ACOG)
Заболевания с тяжёлой клинической картиной
Заболевания, значительно снижающие качество жизни пациентов
Заболевания, развивающиеся в первые годы жизни
Заболевания, для которых доступно хирургическое или медикаментозное лечение
Благодаря современнейшей технологии анализа ДНК с точностью более 99% будущие родители могут пройти самый полный анализ и получить чёткую картину здоровья своего ребёнка.
Гены анализируются методом полного секвенирования гена (full-gene sequencing) с обязательным анализом делеций и дупликаций в значимых областях +/- 10 пар оснований.
Оба метода выполняются по технологии Next generation sequencing (NGS).
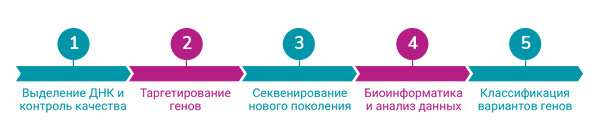
Какие этапы проходит ваш образец крови во время анализа?
Выберите тест, который подходит именно вам.
LIGHT PCR
PCR
 2 гена
● Муковисцидоз, F508del
● Спинальная мышечная атрофия, ген SMN1
160 EUR (≈ 18 800 RSD)
110 EUR (≈ 12 900 RSD)
LIGHT NGS
2 гена
● Спинальная мышечная атрофия, ген SMN1
270 EUR (≈ 31 700 RSD)
цена вместе с тестом Verified
190 EUR (≈ 22 300 RSD)
PLUS
30 генов
список заболеваний
480 EUR (≈ 56 300 RSD)
цена вместе с тестом Verified
400 EUR (≈ 46 900 RSD)
400 генов
список заболеваний
● Полная и точная информация о более чем 450 наиболее значимых моногенных заболеваниях
660 EUR (≈ 77 500 RSD)
цена вместе с тестом Verified
550 EUR (≈ 64 600 RSD)
COMPLETE
2200 генов
список заболеваний
● Комплексный анализ широкого спектра генных мутаций
880 EUR (≈ 103 300 RSD)
750 EUR (≈ 88 000 RSD)
Инфоцентр
+381 11 414 65 65
Заказать тест
Пожалуйста, введите свои данные
Этот сайт защищён reCAPTCHA, и к нему применяются Политика конфиденциальности и Условия использования Google.
Результаты и дальнейшие шаги
Возможные результаты теста ENVISION:
- Изменения в исследованных генах не выявлены
+ В исследованных генах выявлены изменения
Каковы дальнейшие шаги?
Если вы и ваш партнёр — носители изменённого гена, наши генетики проконсультируют вас о точном риске заболевания у малыша и о дальнейших шагах, которые могут включать:
● Диагностические тесты во время беременности — амниоцентез или биопсия ворсин хориона
● Альтернативные методы зачатия
● PGT – M
Список заболеваний, которые анализирует
| ABCA3 | Surfactant metabolism dysfunction, pulmonary, type 3 |
| ABCB11 | Cholestasis, benign recurrent intrahepatic, type 2; Cholestasis, progressive familial intrahepatic, type 2 |
| ABCC8 | Hyperinsulinemic hypoglycemia, type 1 (congenital hyperinsulinism); Permanent neonatal diabetes mellitus (PNDM) |
| ABCD1 | Adrenoleukodystrophy |
| ABCD4 | Methylmalonic aciduria and homocystinuria, cblJ type |
| ACAD9 | Acyl-CoA dehydrogenase 9 deficiency (mitochondrial complex I deficiency, nuclear, type 20) |
| ACADM | Medium-chain acyl-CoA dehydrogenase deficiency |
| ACADS | Short-chain acyl-CoA dehydrogenase deficiency |
| ACADSB | Short/branched-chain acyl-CoA dehydrogenase deficiency (2-methylbuyrylglycinuria) |
| ACADVL | Very long-chain acyl-CoA dehydrogenase (VLCAD) deficiency |
| ACAT 1 | Alpha-methylacetoacetic aciduria (3-ketothiolase deficiency) |
| ACOX1 | Peroxisomal acyl-CoA oxidase deficiency |
| ACSF3 | Combined malonic and methylmalonic aciduria |
| ADA | Severe combined immunodeficiency due to adenosine deaminase deficiency (ADA) |
| ADAMTS2 | Ehlers-Danlos syndrome, dermatosparaxis type |
| ADGRG1 | Polymicrogyria, bilateral frontoparietal |
| ADK | Hypermethioninemia due to adenosine kinase deficiency |
| AFF2 | Intellectual developmental disorder, X-linked 109 |
| AGA | Aspartylglucosaminuria (glycosylasparaginase deficiency) |
| AGL | Glycogen storage disease, type 3 |
| AGPS | Rhizomelic chondrodysplasia punctata, type 3 |
| AGXT | Hyperoxaluria, primary, type 1 |
| AHCY | Hypermethioninemia with deficiency of S-adenosylhomocysteine hydrolase |
| AHI1 | Joubert syndrome, type 3 |
| AIPL1 | Leber congenital amaurosis, type 4 |
| AIRE | Autoimmune polyendocrinopathy syndrome, type 1 |
| ALDH3A2 | Sjogren-Larsson syndrome |
| ALDH4A1 | Hyperprolinemia, type 2 |
| ALDOB | Fructose intolerance, hereditary |
| ALG6 | Congenital disorder of glycosylation, type 1C |
| ALMS1 | Alström syndrome |
| ALPL | Hypophosphatasia, infantile/childhood |
| AMT | Glycine encephalopathy |
| ANO10 | Spinocerebellar ataxia, autosomal recessive, type 10 |
| AP1S2 | Pettigrew syndrome |
| AQP2 | Diabetes insipidus, nephrogenic, type 2 |
| ARG1 | Argininemia (arginase deficiency) |
| ARL13B | Joubert syndrome type 8 |
| ARSA | Metachromatic leukodystrophy |
| ARSB | Mucopolysaccharidosis, type 6 (Maroteaux-Lamy syndrome) |
| ARSE | Chondrodysplasia punctata, X-linked recessive |
| ARX | Epileptic encephalopathy, early infantile, type 1; ARX-related developmental disorders |
| ASL | Argininosuccinic aciduria |
| ASNS | Asparagine synthetase deficiency |
| ASPA | Canavan disease |
| ASS1 | Citrullinemia, type 1 |
| ATM | Ataxia-telangiectasia |
| ATP6V1B1 | Renal tubular acidosis with deafness |
| ATP7A | Menkes disease; Occipital horn syndrome |
| ATP7B | Wilson disease |
| ATRX | Intellectual disability-hypotonic facies syndrome, X-linked; Alpha-thalassemia/intellectual developmental disorder syndrome |
| BBS1 | Bardet-Biedl syndrome, type 1 |
| BBS10 | Bardet-Biedl syndrome, type 10 |
| BBS12 | Bardet-Biedl syndrome, type 12 |
| BBS2 | Bardet-Biedl syndrome, type 2 |
| BCHE | Butyrylcholinesterase deficiency |
| BCKDHA | Maple syrup urine disease, type 1A |
| BCKDHB | Maple syrup urine disease, type 1B |
| BCS1L | Mitochondrial complex III deficiency nuclear type 1; GRACILE syndrome; Bjornstad syndrome |
| BLM | Bloom syndrome |
| BRWD3 | Intellectual developmental disorder, X-linked 93 |
| BSND | Bartter syndrome, type 4A |
| BTD | Biotinidase deficiency |
| CAPN3 | Limb-girdle muscular dystrophy, type 1 (LGMD R1) |
| CASQ2 | Ventricular tachycardia, catecholaminergic polymorphic, type 2 |
| CBS | Homocystinuria due to cystathionine beta-synthase |
| CC2D2A | Joubert syndrome, type 9; Meckel syndrome, type 6; COACH syndrome, 2 |
| CCDC103 | Ciliary dyskinesia, primary, type 17 |
| CCDC151 | Primary ciliary dyskinesia, type 30 |
| CCDC39 | Ciliary dyskinesia, primary, type 14 |
| CCDC88C | Hydrocephalus, congenital, type 1 |
| CD40LG | Hyper-IgM syndrome, type 1 (immunodeficiency, X-linked, with hyper-IgM, type 1) |
| CDH23 | Deafness, autosomal recessive, type 12; Usher syndrome, type 1D |
| CEP290 | Meckel syndrome, type 4; Joubert syndrome, type 5; Leber congenital amaurosis, type 10 |
| CERKL | Retinitis pigmentosa, type 26 |
| CFTR | Cystic fibrosis |
| CHM | Choroideremia |
| CHRNE | Myasthenic syndrome, congenital, type 4B, fast-channel; Myasthenic syndrome, congenital, type 4C, associated with acetylcholine receptor deficiency |
| CHRNG | Multiple pterygium syndrome (MPS), Escobar type; MPS, lethal type |
| CHST6 | Macular corneal dystrophy |
| CIITA | Bare lymphocyte syndrome, type 2 complementation group A |
| CLCN1 | Myotonia congenita, recessive |
| CLN3 | Ceroid lipofuscinosis, neuronal, type 3 |
| CLN5 | Ceroid lipofuscinosis, neuronal, type 5 |
| CLN6 | Ceroid lipofuscinosis, neuronal, type 6 |
| CLN8 | Ceroid lipofuscinosis, neuronal, type 8 |
| CLRN1 | Usher syndrome, type 3A |
| CNGA1 | Retinitis pigmentosa type 49 |
| CNGB1 | Retinitis pigmentosa type 45 |
| CNGB3 | Achromatopsia, type 3 |
| COL 4 A3 | Alport syndrome, autosomal recessive, type 2 |
| COL 4 A4 | Alport syndrome, autosomal recessive, type 2 |
| COL 4 A5 | Alport syndrome, X-linked |
| COL27A1 | Steel syndrome |
| COL7A1 | Dystrophic epidermolysis bullosa (DEB), Hallopeau-Siemens (HS) type and non-HS type; DEB pruriginosa; DEB pretibial |
| CPS1 | Carbamoylphosphate synthetase 1 deficiency |
| CPT1A | Carnitine palmitoyltransferase type 1A deficiency, hepatic |
| CPT2 | Carnitine palmitoyltransferase type 2 deficiency, lethal neonatal; Carnitine palmitoyltransferase type 2 deficiency, infantile |
| CRB1 | Retinitis pigmentosa, type 12; Leber congenital amaurosis, type 8 |
| CTNS | Nephropathic cystinosis |
| CTSK | Pycnodysostosis |
| CUL4B | Intellectual developmental disorder, X-linked syndromic, Cabezas type |
| CYBA | Chronic granulomatous disease, type 4 |
| CYBB | Chronic granulomatous disease, X-linked |
| CYP 11 A1 | 46,XY disorder of sex development-adrenal insufficiency due to CYP11A1 deficiency |
| CYP11B1 | Adrenal hyperplasia, congenital, due to 11-beta-hydroxylase deficiency |
| CYP11B2 | Hypoaldosteronism, congenital, due to CMO I deficiency |
| CYP17A1 | 17 alpha-hydroxylase/17,20-lyase deficiency |
| CYP19A1 | Aromatase deficiency |
| CYP1B1 | Glaucoma, primary congenital, type 3A |
| CYP21A2 | Congenital adrenal hyperplasia due to 21-hydroxylase deficiency |
| CYP27A1 | Cerebrotendinous xanthomatosis |
| CYP27B1 | Vitamin D-dependent rickets, type 1 |
| DBT | Maple syrup urine disease, type 2 |
| DCLRE1C | Omenn syndrome; Severe combined immunodeficiency, Athabascan type |
| DCX | Lissencephaly, X-linked, type 1 |
| DHCR7 | Smith-Lemli-Opitz syndrome |
| DHDDS | Retinitis pigmentosa, type 59 |
| DLD | Dihydrolipoamide dehydrogenase deficiency |
| DLG3 | Intellectual developmental disorder, X-linked 90 |
| DMD | Duchenne/Becker muscular dystrophy |
| DNAH5 | Ciliary dyskinesia, primary, type 3 with or without situs inversus |
| DNAI1 | Ciliary dyskinesia, primary, type 1 with or without situs inversus |
| DNAI2 | Ciliary dyskinesia, primary, type 9 with or without situs inversus |
| DNAL1 | Ciliary dyskinesia, primary, type 16 |
| DPYD | Dihydropyrimidine dehydrogenase deficiency |
| DUOX2 | Thyroid dyshormonogenesis, type 6 |
| DUOXA2 | Thyroid dyshormonogenesis, type 5 |
| DYNC2H1 | Short-rib thoracic dysplasia, type 3 with or without polydactyly |
| DYSF | Miyoshi muscular dystrophy, type 1; Limb-girdle muscular dystrophy, type 2 (LGMD R2) |
| EDA | Ectodermal dysplasia, type 1 hypohidrotic, X-linked |
| EIF2AK3 | Wolcott-Rallison syndrome |
| EIF2B5 | Leukoencephalopathy with vanishing white matter (VWM) |
| ELP1 (IKBKAP) | Familial dysautonomia |
| EMD | Emery-Dreifuss muscular dystrophy, type 1 X-linked |
| ERCC2 | Trichothiodystrophy, type 1 |
| ERCC6 | Cockayne syndrome, type B; Cerebrooculofacioskeletal syndrome, type 1 |
| ERCC8 | Cockayne syndrome, type A |
| ESCO2 | Roberts syndrome; Juberg-Hayward syndrome |
| ETFA | Glutaric acidemia, type 2A |
| ETFB | Glutaric acidemia, type 2B |
| ETFDH | Glutaric acidemia, type 2C |
| ETHE1 | Ethylmalonic encephalopathy |
| EVC | Ellis-van Creveld syndrome |
| EVC2 | Ellis-van Creveld syndrome |
| EXOSC3 | Pontocerebellar hypoplasia, type 1B |
| EYS | Retinitis pigmentosa, type 25 |
| F11 | Factor XI deficiency |
| F2 | Prothrombin deficiency |
| F5 | Factor V deficiency |
| F8 | Hemophilia A |
| F9 | Hemophilia B |
| FAH | Tyrosinemia, type 1 |
| FAM161A | Retinitis pigmentosa, type 28 |
| FANCA | Fanconi anemia, complementation group A |
| FANCC | Fanconi anemia, complementation group C |
| FANCG | Fanconi anemia, complementation group G |
| FGD1 | Aarskog-Scott syndrome; intellectual developmental disorder, X-linked syndromic, type 16 |
| FH | Fumarase deficiency |
| FKRP | Muscular dystrophy-dystroglycanopathy, type 5A (Walker-Warburg syndrome); Type 5B; Type 5C (limb-girdle muscular dystrophy, type 9 [LGMDR9]) |
| FKTN | Muscular dystrophy-dystroglycanopathy, type 4A (Walker-Warburg syndrome); Type 4B; Type 4C (limb-girdle muscular dystrophy, type 13 [LGMD R13]) |
| FMO3 | Trimethylaminuria |
| FMR1 | Fragile X syndrome |
| FTCD | Glutamate formiminotransferase deficiency |
| FTSJ1 | Intellectual developmental disorder, X-linked 9 |
| FXN | Friedreich ataxia |
| G6PC | Glycogen storage disease, type 1A |
| G6PD | Hemolytic anemia, G6PD deficient (favism) |
| GAA | Glycogen storage disease, type 2 |
| GALC | Krabbe disease |
| GALE | Galactose epimerase deficiency |
| GALK1 | Galactokinase deficiency with cataracts |
| GALNS | Mucopolysaccharidosis, type 4A |
| GALT | Galactosemia |
| GAMT | Cerebral creatine deficiency syndrome, type 2 |
| GBA | Gaucher Disease |
| GBE1 | Glycogen storage disease, type 4 |
| GCDH | Glutaricaciduria, type 1 |
| GDAP1 | Charcot-Marie-Tooth disease, recessive intermediate, type A |
| GFM1 | Combined oxidative phosphorylation deficiency, type 1 |
| GJB1 | Charcot-Marie-Tooth neuropathy, X-linked dominant, type 1 |
| GJB2 | Deafness, autosomal recessive, type 1A; Deafness, digenic, GJB2/GJB6 |
| GJB6 | Deafness, autosomal recessive, type 1B; Deafness, digenic GJB2/GJB6 |
| GLA | Fabry disease |
| GLB1 | GM1-gangliosidosis, types 1-3; Mucopolysaccharidosis, type 4B (Morquio) |
| GLDC | Glycine encephalopathy |
| GLE1 | Lethal congenital contracture syndrome, type 1; Congenital arthrogryposis with anterior horn cell disease |
| GNE | Inclusion body myopathy, type 2 (Nonaka myopathy) |
| GNPTAB | Mucolipidosis 2 alpha/beta; Mucolipidosis 3 alpha/beta |
| GNPTG | Mucolipidosis III gamma |
| GNRHR | Hypogonadotropic hypogonadism, type 7 without anosmia |
| GNS | Mucopolysaccharidosis, type 3D (Sanfilippo syndrome D) |
| GP1BA | Bernard-Soulier syndrome, type A1 |
| GP9 | Bernard-Soulier syndrome, type C |
| GPR143 | Ocular albinism, type 1 (Nettleship-Falls type) |
| GRHPR | Hyperoxaluria, primary, type 2 |
| GRIP1 | Fraser syndrome 3 |
| GUSB | Mucopolysaccharidosis, type 7 |
| HADHA | Long-chain 3-hydroxyl-CoA dehydrogenase (LCHAD) deficiency; Mitochondrial trifunctional protein deficiency |
| HAX1 | Neutropenia, severe congenital, type 3 autosomal recessive |
| HBA1 / HBA2 | Alpha thalassemia |
| HBB | Beta-thalassemia, Sickle cell anemia and other HBB-related hemoglobinopathies |
| HEXA | Tay-Sachs disease |
| HEXB | Sandhoff disease, infantile, juvenile, and adult forms |
| HFE | Hemochromatosis |
| HGD | Alkaptonuria |
| HGSNAT | Mucopolysaccharidosis type 3C (Sanfilippo syndrome C) |
| HJV (HFE2) | Hemochromatosis, type 2A |
| HLCS | Holocarboxylase synthetase deficiency |
| HMGCL | HMG-CoA lyase deficiency |
| HOGA1 | Hyperoxaluria, primary, type 3 |
| HPS1 | Hermansky-Pudlak syndrome, type 1 |
| HPS3 | Hermansky-Pudlak syndrome, type 3 |
| HSD17B4 | D-bifunctional protein deficiency |
| HSD3B2 | Adrenal hyperplasia, congenital, due to 3-beta-hydroxysteroid dehydrogenase 2 deficiency |
| HYAL1 | ?Mucopolysaccharidosis, type 9 |
| HYLS1 | Hydrolethalus syndrome |
| IDH3B | Retinitis pigmentosa, type 46 |
| IDS | Mucopolysaccharidosis, type 2 |
| IDUA | Mucopolysaccharidosis type 1 |
| IL1RAPL1 | Intellectual developmental disorder, X-linked 21 |
| IL2RG | Severe combined immunodeficiency, X-linked |
| IVD | Isovaleric acidemia |
| IYD | Thyroid dyshormonogenesis, type 4 |
| JAK3 | Severe Combined Immunodeficiency, autosomal recessive, T-negative/B-positive type |
| KCNJ11 | Hyperinsulinemic hypoglycemia, type 2 (congenital hyperinsulinism); Permanent neonatal diabetes mellitus (PNDM) |
| KDM5C | Intellectual developmental disorder, X-linked syndromic, Claes-Jensen type |
| L1CAM | L1 Syndrome |
| LAMA2 | LAMA2-related muscular dystrophy |
| LAMA3 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LAMB3 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LAMC2 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LCA5 | Leber congenital amaurosis, type 5 |
| LDLRAP1 | Hypercholesterolemia, familial, autosomal recessive |
| LHX3 | Pituitary hormone deficiency, combined, type 3 |
| LIFR | Stuve-Wiedemann syndrome / Schwartz-Jampel type 2 syndrome |
| LIPA | Lysosomal Acid Lipase Deficiency |
| LMBRD1 | Methylmalonic aciduria and homocystinuria, cblF type |
| LOXHD1 | Deafness, autosomal recessive, type 77 |
| LPL | Lipoprotein lipase deficiency |
| LRP2 | Donnai-Barrow syndrome |
| LRPPRC | Mitochondrial complex IV deficiency, nuclear type 5, (French-Canadian) |
| LYST | Chediak-Higashi syndrome |
| MAN2B1 | Alpha-mannosidosis |
| MCCC1 | 3-Methylcrotonyl-CoA carboxylase deficiency, type 1 |
| MCCC2 | 3-Methylcrotonyl-CoA carboxylase deficiency, type 2 |
| MCEE | Methylmalonyl-CoA epimerase deficiency |
| MCOLN1 | Mucolipidosis type 4 |
| MCPH1 | Microcephaly type 1 primary, autosomal recessive |
| MED17 | Microcephaly, postnatal progressive, with seizures and brain atrophy |
| MEFV | Familial Mediterranean fever |
| MESP2 | Spondylocostal dysostosis, type 2 autosomal recessive |
| MFSD8 | Ceroid lipofuscinosis, neuronal, type 7 |
| MID1 | Opitz GBBB syndrome, type 1 |
| MKS1 | Bardet-Biedl syndrome type 13; Meckel syndrome, type 1; Joubert syndrome, type 28 |
| MLC1 | Megalencephalic leukoencephalopathy with subcortical cysts |
| MMAA | Methylmalonic aciduria, vitamin B12-responsive |
| MMAB | Methylmalonic aciduria, vitamin B12-responsive, type cblB |
| MMACHC | Methylmalonic aciduria and homocystinuria, cblC type |
| MMADHC | Homocystinuria, cblD type, variant 1 |
| MPI | Congenital disorder of glycosylation, type 1B |
| MPL | Thrombocytopenia, congenital amegakaryocytic |
| MPV17 | Mitochondrial DNA depletion syndrome type 6 (hepatocerebral); Charcot-Marie-Tooth disease, axonal, type 2EE |
| MTHFR | Homocystinuria due to MTHFR deficiency |
| MTM1 | Myotubular myopathy, X-linked |
| MTMR2 | Charcot-Marie-Tooth disease, type 4B1 |
| MTRR | Homocystinuria-megaloblastic anemia, cbl E type |
| MTTP | Abetalipoproteinemia |
| MUT | Methylmalonic acidemia (MUT-related) |
| MVK | Mevalonic aciduria |
| MYO7A | Usher syndrome, type 1B; Deafness, autosomal recessive, type 2 |
| NAGA | Schindler disease, type I |
| NAGLU | Mucopolysaccharidosis, type 3B (Sanfilippo B) |
| NAGS | N-acetylglutamate synthase deficiency |
| NBN | Nijmegen breakage syndrome |
| NDP | Norrie disease |
| NDRG1 | Charcot-Marie-Tooth disease, type 4D |
| NDUFAF5 | Mitochondrial complex I deficiency, nuclear type 16 |
| NDUFS6 | Mitochondrial complex I deficiency, nuclear type 9 |
| NEB | Nemaline myopathy type 2 |
| NPC1 | Niemann-Pick disease, type C1 |
| NPC2 | Niemann-pick disease, type C2 |
| NPHP1 | Joubert syndrome type 4 |
| NPHS1 | Nephrotic syndrome, type 1 |
| NPHS2 | Nephrotic syndrome, type 2 |
| NR0B1 | Adrenal hypoplasia, congenital |
| NR2E3 | Enhanced S-cone syndrome (Goldmann-Favre); Retinitis pigmentosa, type 37 |
| NTRK1 | Insensitivity to pain, congenital, with anhidrosis |
| OAT | Gyrate atrophy of choroid and retina |
| OCA2 | Oculocutaneous albinism type 2 |
| OCRL | Lowe Syndrome; Dent disease type 2 |
| OPA3 | 3-methylglutaconic aciduria, type 3 |
| OPHN1 | Intellectual developmental disorder, X-linked syndromic, Billuart type |
| OTC | Ornithine transcarbamylase deficiency |
| OTOF | Deafness, autosomal recessive, type 9 |
| P3H1 | Osteogenesis imperfecta, type 8 |
| PAH | Phenylketonuria |
| PAK3 | Intellectual developmental disorder, X-linked 30 |
| PANK2 | Neurodegeneration with brain iron accumulation type 1 |
| PC | Pyruvate carboxylase deficiency |
| PCBD1 | Hyperphenylalaninemia, BH4-deficient, type D |
| PCCA | Propionic acidemia |
| PCCB | Propionic acidemia |
| PCDH15 | Deafness, autosomal recessive, type 23; Usher syndrome, type 1D/F digenic |
| PDE6A | Retinitis pigmentosa type 43 |
| PDHA1 | Pyruvate dehydrogenase E1-alpha deficiency |
| PDHB | Pyruvate dehydrogenase E1-beta deficiency |
| PEX1 | Heimler syndrome type 1 |
| PEX10 | Peroxisome biogenesis disorder, type 6A (Zellweger syndrome); Peroxisome biogenesis disorder, type 6B |
| PEX12 | Peroxisome biogenesis disorder type 3A (Zellweger) |
| PEX2 | Peroxisome biogenesis disorder type 5A (Zellweger) |
| PEX6 | Peroxisome biogenesis disorder, type 4A (Zellweger syndrome); Peroxisome biogenesis disorder, type 4B; Heimler syndrome 2 |
| PEX7 | Rhizomelic chondrodysplasia punctata, type 1 |
| PFKM | Glycogen storage disease, type 7 |
| PGK1 | Phosphoglycerate kinase 1 deficiency |
| PHF8 | Intellectual developmental disorder, X-linked syndromic, Siderius type |
| PHGDH | Neu-Laxova syndrome, type 1; Phosphoglycerate dehydrogenase deficiency |
| PKHD1 | Polycystic kidney disease type 4 |
| PLA2G6 | Infantile neuroaxonal dystrophy type 1 |
| PLOD1 | Ehlers-Danlos syndrome, kyphoscoliotic type, 1 |
| PLP1 | Pelizaeus-Merzbacher disease |
| PMM2 | Congenital disorder of glycosylation, type 1A |
| POLG | POLG-related disorders |
| POLR1C | Leukodystrophy, hypomyelinating, type 11; Treacher Collins syndrome 3 |
| POMGNT1 | Muscular dystrophy-dystroglycanopathy, type 3A (Walker-Warburg syndrome); Type 3B; Type 3C (limb-girdle muscular dystrophy, type 15 [LGMDR15]) |
| POMT1 | Muscular dystrophy-dystroglycanopathy, type 1A (Walker-Warburg syndrome); Type 1B; Type 1C (limb-girdle muscular dystrophy, type 11 [LGMD R11]) muscular dystrophy, type 11 [LGMD R11]) |
| POMT2 | Muscular dystrophy-dystroglycanopathy, type 2A (Walker-Warburg syndrome); Type 2B; Type 2C (limb-girdle muscular dystrophy, type 14 [LGMD R14]) muscular dystrophy, type 14 [LGMD R14]) |
| POU3F4 | Deafness, X-linked, type 2 |
| PPT1 | Ceroid lipofuscinosis, neuronal, type 1 |
| PQBP1 | Renpenning syndrome |
| PRF1 | Hemophagocytic lymphohistiocytosis, familial, type 2 |
| PROP1 | Pituitary hormone deficiency, combined, type 2 |
| PRPS1 | Phosphoribosylpyrophosphate synthetase (PRS) deficiency |
| PSAP | Combined SAP deficiency |
| PTS | Hyperphenylalaninemia, BH4-deficient, type A |
| PUS1 | Myopathy, lactic acidosis, and sideroblastic anemia, type 1 |
| PYGM | McArdle disease |
| QDPR | Hyperphenylalaninemia, BH4-deficient, type C |
| RAB23 | Carpenter syndrome |
| RAG1 | Omenn syndrome; Severe combined immunodeficiency, B cell-negative |
| RAG2 | Omenn syndrome; Severe combined immunodeficiency, B cell-negative |
| RAPSN | Fetal akinesia deformation sequence, type 2; Myasthenic syndrome, congenital, type 11 associated with AChR deficiency |
| RARS2 | Pontocerebellar hypoplasia, type 6 |
| RAX | Microphthalmia, syndromic 16 |
| RDH12 | Leber congenital amaurosis, type 13 |
| RMRP | Anauxetic dysplasia 1 |
| RNASEH2B | Aicardi-Goutieres syndrome, type 2 |
| RP2 | Retinitis pigmentosa, type 2 X-linked |
| RPE65 | RPE65-related Leber congenital amaurosis/early-onset severe retinal dystrophy |
| RPGR | Retinitis pigmentosa, type 3 X-linked; Cone-rod dystrophy, X-linked, 1 |
| RPGRIP1L | Joubert syndrome, type 7; Meckel syndrome, type 5; COACH syndrome |
| RS1 | Retinoschisis |
| RTEL1 | Dyskeratosis congenita, autosomal recessive type 5 |
| SACS | Spastic ataxia, Charlevoix-Saguenay, type |
| SAMHD1 | Aicardi-Goutieres syndrome, type 5 |
| SCO2 | Mitochondrial complex IV deficiency, nuclear type 2 |
| SEPSECS | Pontocerebellar hypoplasia, type 2D |
| SERPINA1 | Alpha-1 antitrypsin deficiency |
| SGCA | Limb-girdle muscular dystrophy, type 3 (LGMD R3) |
| SGCB | Limb-girdle muscular dystrophy, type 4 (LGMD R4) |
| SGCD | Limb-girdle muscular dystrophy, type 6 (LGMD R6) |
| SGCG | Limb-girdle muscular dystrophy, type 5 (LGMD R5) |
| SGSH | Mucopolysaccharidosis, type 3A (Sanfilippo A) |
| SH3TC2 | Charcot-Marie-Tooth disease, type 4C |
| SLC 39A4 | Acrodermatitis enteropathica |
| SLC12 A6 | Agenesis of the corpus callosum with peripheral neuropathy |
| SLC12A3 | Gitelman syndrome |
| SLC16 A2 | Allan-Herndon-Dudley syndrome |
| SLC17A5 | Salla disease |
| SLC19A3 | Thiamine metabolism dysfunction syndrome, type 2 (biotin- or thiamine-responsive encephalopathy type) |
| SLC22A5 | Carnitine deficiency, systemic primary |
| SLC25A13 | Citrullinemia, type 2 neonatal-onset; Citrullinemia, type 2 adult-onset |
| SLC25A15 | Hyperornithinemia-hyperammonemia-homocitrullinemia syndrome |
| SLC25A20 | Carnitine-acylcarnitine translocase deficiency |
| SLC26 A2 | Achondrogenesis, type 1B (diastrophic dysplasia) |
| SLC26A3 | Diarrhea 1 secretory chloride, congenital |
| SLC26A4 | Deafness, autosomal recessive, type 4; Pendred syndrome |
| SLC35A3 | Arthrogryposis, impaired intellectual development, and seizures |
| SLC37A4 | Glycogen storage disease, type 1B |
| SLC46A1 | Folate malabsorption, hereditary |
| SLC4A11 | Corneal endothelial dystrophy, autosomal recessive |
| SLC5A5 | Thyroid dyshormonogenesis, type 1 |
| SLC6A19 | Hartnup disorder |
| SLC6A8 | Cerebral creatine deficiency syndrome, type 1 |
| SLC7A7 | Lysinuric protein intolerance |
| SMARCAL1 | Schimke immunoosseous dysplasia |
| SMN1 | Spinal muscular atrophy |
| SMPD1 | Niemann-Pick disease, type A; Niemann-Pick disease, type B |
| SPG11 | Amyotrophic lateral sclerosis, type 5 juvenile |
| SPG7 | Spastic paraplegia, type 7 autosomal recessive |
| STAR | Lipoid adrenal hyperplasia |
| SUMF1 | Multiple sulfatase deficiency |
| SURF1 | Charcot-Marie-Tooth disease, type 4K; Leigh syndrome, due to COX IV deficiency |
| SYN1 | Epilepsy, X-linked, with variable learning disabilities and behavior disorders |
| TAT | Tyrosinemia, type 2 |
| TCIRG1 | Osteopetrosis, autosomal recessive, type 1 |
| TECPR2 | Spastic paraplegia, type 49 autosomal recessive |
| TF | Atransferrinemia |
| TFR2 | Hemochromatosis, type 3 |
| TG | Thyroid dyshormonogenesis, type 3 |
| TGM1 | Ichthyosis, congenital, autosomal recessive, type 1 |
| TH | Segawa syndrome, recessive |
| THOC2 | Intellectual developmental disorder, X-linked 12 |
| TMEM216 | Joubert syndrome, type 2; Meckel syndrome, type 2 |
| TNXB | Ehlers-Danlos syndrome, classic-like |
| TPO | Thyroid dyshormonogenesis, type 2A |
| TPP1 | Ceroid lipofuscinosis, neuronal, type 2; Spinocerebellar ataxia, autosomal recessive, type 7 |
| TRDN | Ventricular tachycardia, catecholaminergic polymorphic, type 5 with or without muscle weakness |
| TRIM32 | Limb-girdle muscular dystrophy, type 8 (LGMD R8) |
| TRMU | Liver failure, transient infantile |
| TSFM | Combined oxidative phosphorylation deficiency, type 3 |
| TSHB | Hypothyroidism, congenital, nongoitrous, type 4 |
| TTC37 | Trichohepatoenteric syndrome 1 |
| TTPA | Ataxia with isolated vitamin E deficiency |
| TYMP | Mitochondrial DNA depletion syndrome, type 1 (MNGIE type) |
| TYR | Oculocutaneous albinism (OCA) type 1A; OCA type 1B |
| UGT1A1 | Crigler-Najjar syndrome, type 1; Crigler-Najjar syndrome, type 2 |
| UPF3B | Intellectual developmental disorder, X-linked syndromic 14 |
| USH1C | Usher syndrome, type 1C; Deafness, autosomal recessive, type 18A |
| USH1G | Usher syndrome, type 1G |
| USH2A | Usher syndrome, type 2A |
| VPS13A | Choreoacanthocytosis |
| VPS13B | Cohen syndrome |
| VPS45 | Neutropenia, severe congenital, type 5 |
| VRK1 | Pontocerebellar hypoplasia, type 1A |
| VSX2 | Microphthalmia with coloboma 3; Isolated microphthalmia 2 |
| WAS | Wiskott-Aldrich syndrome; Thrombocytopenia, X-linked |
| WHRN | Usher syndrome, type 2D; Deafness, autosomal recessive, type 31 |
| WNT10A | Ectodermal dysplasia 16 (odontoonychodermal dysplasia) |
| XPA | Xeroderma pigmentosum, group A |
| XPC | Xeroderma pigmentosum, group C |
| ZDHHC9 | Intellectual developmental disorder, X-linked syndromic, Raymond type |
| ZFYVE26 | Spastic paraplegia, type 15 autosomal recessive |
| ZNF711 | Intellectual developmental disorder, X-linked 97 |
Список заболеваний, которые анализирует
COMPLETE
| AAAS | Triple-A syndrome (achalasia-addisonianism-alacrimia) |
| AARS1 | Epileptic encephalopathy, early infantile, type 29 |
| AARS2 | Combined oxidative phosphorylation deficiency 8; Leukoencephalopathy, progressive, with ovarian failure |
| AASS | Hyperlysinemia, type 1 and type 2 |
| ABAT | GABA-transaminase deficiency |
| ABCA1 | Tangier disease |
| ABCA12 | Ichthyosis congenital autosomal recessive type 4A and 4B (harlequin) |
| ABCA3 | Surfactant metabolism dysfunction, pulmonary, type 3 |
| ABCA4 | Stargardt disease type 1; Cone-rod dystrophy type 3 |
| ABCB11 | Cholestasis, benign recurrent intrahepatic, type 2; Cholestasis, progressive familial intrahepatic, type 2 |
| ABCB4 | Cholestasis, progressive familial intrahepatic, type 3 |
| ABCB7 | Anemia, sideroblastic, with ataxia |
| ABCC2 | Dubin-Johnson syndrome |
| ABCC6 | Pseudoxanthoma elasticum; Generalized arterial calcification of infancy, type 2 |
| ABCC8 | Hyperinsulinemic hypoglycemia, type 1 (congenital hyperinsulinism); Permanent neonatal diabetes mellitus (PNDM) |
| ABCD1 | Adrenoleukodystrophy |
| ABCD4 | Methylmalonic aciduria and homocystinuria, cblJ type |
| ABCG5 | Sitosterolemia |
| ABCG8 | Sitosterolemia |
| ABHD12 | PHARC syndrome (polyneuropathy, hearing loss, ataxia, retinitis pigmentosa and cataract) |
| ABHD5 | Chanarin-Dorfman syndrome |
| ACAD8 | Isobutyryl-CoA dehydrogenase deficiency |
| ACAD9 | Acyl-CoA dehydrogenase 9 deficiency (mitochondrial complex I deficiency, nuclear, type 20) |
| ACADM | Medium-chain acyl-CoA dehydrogenase deficiency |
| ACADS | Short-chain acyl-CoA dehydrogenase deficiency |
| ACADSB | Short/branched-chain acyl-CoA dehydrogenase deficiency (2-methylbuyrylglycinuria) |
| ACADVL | Very long-chain acyl-CoA dehydrogenase (VLCAD) deficiency |
| ACAT1 | Alpha-methylacetoacetic aciduria (3-ketothiolase deficiency) |
| ACE | Renal tubular dysgenesis |
| ACO2 | Infantile cerebellar-retinal degeneration |
| ACOX1 | Peroxisomal acyl-CoA oxidase deficiency |
| ACOX2 | Bile acid synthesis defect, congenital, type 6 |
| ACP5 | Spondyloenchondrodysplasia with immune dysregulation |
| ACSF3 | Combined malonic and methylmalonic aciduria |
| ACSL4 | Intellectual developmental disorder, X-linked 63 |
| ACTA1 | Nemaline myopathy 3; Congenital fiber-type disproportion myopathy 1 |
| ACTN4 | Glomerulosclerosis, focal segmental, 1 |
| ACY1 | Aminoacylase 1 deficiency |
| ADA | Severe combined immunodeficiency due to adenosine deaminase deficiency (ADA) |
| ADA2 | Vasculitis, autoinflammation, immunodeficiency, and hematologic defects syndrome |
| ADAM9 | Cone-rod dystrophy 9 |
| ADAMTS10 | Weill-Marchesani syndrome, type 1 recessive |
| ADAMTS13 | Thrombotic thrombocytopenic purpura, familial (Schulman-Upshaw syndrome) |
| ADAMTS17 | Weill-Marchesani syndrome, type 4 recessive |
| ADAMTS18 | Microcornea, myopic chorioretinal atrophy, and telecanthus |
| ADAMTS2 | Ehlers-Danlos syndrome, dermatosparaxis type |
| ADAMTSL2 | Geleophysic dysplasia type 1 |
| ADAMTSL4 | Ectopia lentis et pupillae; Ectopia lentis, isolated, type 2 |
| ADAR | Aicardi-Goutieres syndrome, type 6 |
| ADAT3 | Neurodevelopmental disorder with brain abnormalities, poor growth, and dysmorphic facies |
| ADCK3 | Coenzyme Q10 deficiency, primary, 4 |
| ADGRG1 | Polymicrogyria, bilateral frontoparietal |
| ADGRG6 | Lethal congenital contracture syndrome 9 |
| ADGRV1 | Usher syndrome, type 2C |
| ADK | Hypermethioninemia due to adenosine kinase deficiency |
| ADSL | Adenylosuccinase deficiency |
| ADSS1 | Myopathy, distal, 5 |
| AFF2 | Intellectual developmental disorder, X-linked 109 |
| AFG3L2 | Spastic ataxia, type 5 autosomal recessive |
| AFP | Alpha-fetoprotein deficiency |
| AGA | Aspartylglucosaminuria (glycosylasparaginase deficiency) |
| AGBL5 | Retinitis pigmentosa 75 |
| AGK | Cataract 38; Sengers syndrome |
| AGL | Glycogen storage disease, type 3 |
| AGPAT2 | Congenital generalized lipodystrophy (Berardinelli-Seip syndrome) |
| AGPS | Rhizomelic chondrodysplasia punctata, type 3 |
| AGRN | Myasthenic syndrome, congenital, type 8 |
| AGT | Renal tubular dysgenesis |
| AGTR1 | Renal tubular dysgenesis |
| AGXT | Hyperoxaluria, primary, type 1 |
| AHCY | Hypermethioninemia with deficiency of S-adenosylhomocysteine hydrolase |
| AHI1 | Joubert syndrome, type 3 |
| AICDA | Immunodeficiency with hyper-IgM, type 2 |
| AIMP1 | Leukodystrophy, hypomyelinating, type 3 |
| AIMP2 | Leukodystrophy, hypomyelinating, type 17 |
| AIPL1 | Leber congenital amaurosis, type 4 |
| AIRE | Autoimmune polyendocrinopathy syndrome, type 1 |
| AK1 | Hemolytic anemia due to adenylate kinase deficiency |
| AK2 | Reticular dysgenesis |
| AKR1C2 | 46,XY disorder of sex development due to testicular 17,20-desmolase deficiency |
| AKR1D1 | Bile acid synthesis defect, congenital, type 2 |
| ALAD | Porphyria, acute hepatic |
| ALAS2 | Anemia, sideroblastic, 1 |
| ALB | Analbuminemia |
| ALDH18A1 | Spastic paraplegia, type 9B, autosomal recessive; Cutis laxa, type 3A (De Barsy syndrome) |
| ALDH1A3 | Microphthalmia, isolated 8 |
| ALDH3A2 | Sjogren-Larsson syndrome |
| ALDH4A1 | Hyperprolinemia, type 2 |
| ALDH5A1 | Succinic semialdehyde dehydrogenase deficiency |
| ALDH6A1 | Methylmalonate semialdehyde dehydrogenase deficiency |
| ALDH7A1 | Epilepsy, pyridoxine-dependent |
| ALDOA | Glycogen storage disease type 12 |
| ALDOB | Fructose intolerance, hereditary |
| ALG1 | Congenital disorder of glycosylation, type 1K |
| ALG11 | Congenital disorder of glycosylation, type 1P |
| ALG12 | Congenital disorder of glycosylation, type 1G |
| ALG2 | Myasthenic syndrome, congenital, type 14 with tubular aggregates |
| ALG3 | Congenital disorder of glycosylation, type 1D |
| ALG6 | Congenital disorder of glycosylation, type 1C |
| ALG8 | Congenital disorder of glycosylation, type 1H |
| ALG9 | Congenital disorder of glycosylation, type 1L; Gillessen-Kaesbach-Nishimura syndrome |
| ALMS1 | Alström syndrome |
| ALOX12B | Ichthyosis, congenital, autosomal recessive, type 2 |
| ALOXE3 | Ichthyosis, congenital, autosomal recessive, type 3 |
| ALPK3 | Cardiomyopathy, familial hypertrophic, type 27 |
| ALPL | Hypophosphatasia, infantile/childhood |
| ALS2 | Amyotrophic lateral sclerosis, type 2 juvenile; Primary lateral sclerosis, juvenile; Spastic paralysis, infantile onset ascending |
| ALX1 | Frontonasal dysplasia, type 3 |
| ALX3 | Frontonasal dysplasia, type 1 |
| ALX4 | Frontonasal dysplasia, type 2 |
| AMACR | Bile acid synthesis defect, congenital, type 4; Alpha-methylacyl-CoA racemase deficiency |
| AMBN | Amelogenesis imperfecta, type IF |
| AMH | Persistent Mullerian duct syndrome, type 1 |
| AMHR2 | Persistent Mullerian duct syndrome, type II |
| AMN | Megaloblastic anemia 1 (Imerslund-Grasbeck syndrome) |
| AMPD1 | Myopathy due to myoadenylate deaminase deficiency |
| AMPD2 | Pontocerebellar hypoplasia, type 9 |
| AMT | Glycine encephalopathy |
| ANGPTL3 | Hypobetalipoproteinemia, familial, type 2 |
| ANKS6 | Nephronophthisis 16 |
| ANO10 | Spinocerebellar ataxia, autosomal recessive, type 10 |
| ANO5 | Limb-girdle muscular dystrophy, type 12 (LGMD R12) |
| ANTXR1 | GAPO syndrome |
| ANTXR2 | Hyaline fibromatosis syndrome |
| AP1S1 | MEDNIK syndrome |
| AP1S2 | Pettigrew syndrome |
| AP3B1 | Hermansky-Pudlak syndrome, type 2 |
| AP3B2 | Epileptic encephalopathy, early infantile, type 48 |
| AP3D1 | Hermansky-Pudlak syndrome, type 10 |
| AP4B1 | Spastic paraplegia, type 47 autosomal recessive |
| AP4E1 | Spastic paraplegia, type 51 autosomal recessive |
| AP4M1 | Spastic paraplegia, type 50 autosomal recessive |
| AP4S1 | Spastic paraplegia, type 52 autosomal recessive |
| AP5Z1 | Spastic paraplegia, type 48 autosomal recessive |
| APOC2 | Hyperlipoproteinemia, type 1B |
| APOE | Sea-blue histiocyte disease |
| APRT | Adenine phosphoribosyltransferase deficiency |
| APTX | Ataxia, early-onset, with oculomotor apraxia and hypoalbuminemia |
| AQP2 | Diabetes insipidus, nephrogenic, type 2 |
| AR | Androgen insensitivity syndrome, complete |
| ARFGEF2 | Periventricular heterotopia with microcephaly |
| ARG1 | Argininemia (arginase deficiency) |
| ARHGDIA | Nephrotic syndrome, type 8 |
| ARHGEF18 | Retinitis pigmentosa 78 |
| ARHGEF9 | Developmental and epileptic encephalopathy 8 |
| ARL13B | Joubert syndrome type 8 |
| ARL2BP | Retinitis pigmentosa with or without situs inversus |
| ARL6 | Bardet-Biedl syndrome, type 3 |
| ARMC9 | Joubert syndrome 30 |
| ARPC1B | Immunodeficiency, type 71 with inflammatory disease and congenita thrombocytopenia |
| ARSA | Metachromatic leukodystrophy |
| ARSB | Mucopolysaccharidosis, type 6 (Maroteaux-Lamy syndrome) |
| ARSE | Chondrodysplasia punctata, X-linked recessive |
| ARSL | Chondrodysplasia punctata, brachytelephalangic |
| ARV1 | Epileptic encephalopathy, early infantile, 38 |
| ARX | Epileptic encephalopathy, early infantile, type 1; ARX-related developmental disorders |
| ASAH1 | Farber lipogranulomatosis; Spinal muscular atrophy with progressive myoclonic epilepsy |
| ASL | Argininosuccinic aciduria |
| ASNS | Asparagine synthetase deficiency |
| ASPA | Canavan disease |
| ASPH | Traboulsi syndrome |
| ASPM | Primary microcephaly type 5 autosomal recessive |
| ASS1 | Citrullinemia, type 1 |
| ATAD1 | Hyperekplexia 4 |
| ATF6 | Achromatopsia, type 7 |
| ATIC | AICA-ribosiduria due to ATIC deficiency |
| ATM | Ataxia-telangiectasia |
| ATOH7 | Persistent hyperplastic primary vitreous, autosomal recessive |
| ATP13A2 | Kufor-Rakeb syndrome; Spastic paraplegia, type 78 autosomal recessive |
| ATP2A1 | Brody myopathy |
| ATP6AP2 | Parkinsonism with spasticity, X-linked |
| ATP6V0A2 | Cutis laxa, autosomal recessive, type 2A; Wrinkly skin syndrome |
| ATP6V0A4 | Renal tubular acidosis, distal, autosomal recessive |
| ATP6V1A | Cutis laxa, autosomal recessive, type 2D |
| ATP6V1B1 | Renal tubular acidosis with deafness |
| ATP6V1E1 | Cutis laxa, autosomal recessive, type 2C |
| ATP7A | Menkes disease; Occipital horn syndrome |
| ATP7B | Wilson disease |
| ATP8B1 | Cholestasis, progressive familial intrahepatic, type 1; Cholestasis, benign recurrent intrahepatic, type 1 |
| ATR | Seckel syndrome, type 1 |
| ATRX | Intellectual disability-hypotonic facies syndrome, X-linked; Alpha-thalassemia/intellectual developmental disorder syndrome |
| AUH | 3-methylglutaconic aciduria, type 1 |
| AURKC | Spermatogenic failure, type 5 |
| AVIL | Nephrotic syndrome, type 21 |
| AVPR2 | Diabetes insipidus, nephrogenic, 1; Nephrogenic syndrome of inappropriate antidiuresis |
| B2M | Immunodeficiency, type 43 |
| B3GALNT2 | Muscular dystrophy-dystroglycanopathy (congenital with brain and eye anomalies, type A, 11 |
| B3GALT6 | Ehlers-Danlos syndrome, spondylodysplastic type, 2 |
| B3GAT3 | Multiple joint dislocations, short stature, craniofacial dysmorphism, with or without congenital heart defects |
| B3GLCT | Peters-plus syndrome |
| B4GALNT1 | Spastic paraplegia, type 26 autosomal recessive |
| B4GALT1 | Congenital disorder of glycosylation, type 2D |
| B4GALT7 | Ehlers-Danlos syndrome, spondylodysplastic, type 1 |
| B4GAT1 | Muscular dystrophy-dystroglycanopathy (congenital with brain and eye anomalies), type A, 13 |
| B9D1 | Joubert syndrome, type 27; ?Meckel syndrome 9 |
| B9D2 | Joubert syndrome, type 34; ?Meckel syndrome, type 10 |
| BBS1 | Bardet-Biedl syndrome, type 1 |
| BBS10 | Bardet-Biedl syndrome, type 10 |
| BBS12 | Bardet-Biedl syndrome, type 12 |
| BBS2 | Bardet-Biedl syndrome, type 2 |
| BBS4 | Bardet-Biedl syndrome, type 4 |
| BBS5 | Bardet-Biedl syndrome, type 5 |
| BBS7 | Bardet-Biedl syndrome, type 7 |
| BBS9 | Bardet-Biedl syndrome, type 9 |
| BCAT2 | Hypervalinemia or hyperleucine-isoleucinemia |
| BCHE | Butyrylcholinesterase deficiency |
| BCKDHA | Maple syrup urine disease, type 1A |
| BCKDHB | Maple syrup urine disease, type 1B |
| BCKDK | Branched-chain ketoacid dehydrogenase kinase deficiency |
| BCL10 | Immunodeficiency, type 37 |
| BCOR | Microphthalmia, syndromic 2 |
| BCS1L | Mitochondrial complex III deficiency nuclear type 1; GRACILE syndrome; Bjornstad syndrome |
| BEST1 | Bestrophinopathy, AR |
| BFSP1 | Cataract 33 multiple types |
| BHLHA9 | Syndactyly, mesoaxial synostotic, with phalangeal reduction |
| BIN1 | Centronuclear myopathy, type 2 |
| BLM | Bloom syndrome |
| BLNK | Agammaglobulinemia 4 |
| BLOC1S3 | Hermansky-Pudlak syndrome, type 8 |
| BLOC1S6 | Hermansky-Pudlak syndrome, type 9 |
| BLVRA | Hyperbiliverdinemia |
| BMP1 | Osteogenesis imperfecta, type 13 |
| BMPER | Diaphanospondylodysostosis |
| BMPR1B | Acromesomelic dysplasia, Demirhan type |
| BOLA3 | Multiple mitochondrial dysfunctions syndrome 2 with hyperglycinemia |
| BPGM | Erythrocytosis due to bisphosphoglycerate mutase deficiency |
| BPNT2 | Chondrodysplasia with joint dislocations, GPAPP type |
| BRAT1 | Rigidity and multifocal seizure syndrome, lethal neonatal; Neurodevelopmental disorder with cerebellar atrophy and with or without seizures |
| BRCA2 | Fanconi anemia, complementation group D1 |
| BRF1 | Cerebellofaciodental syndrome |
| BRIP1 | Fanconi anemia, complementation group J |
| BRWD3 | Intellectual developmental disorder, X-linked 93 |
| BSCL2 | Congenital generalized lipodystrophy, type 2; Encephalopathy, progressive, with or without lipodystrophy |
| BSND | Bartter syndrome, type 4A |
| BTD | Biotinidase deficiency |
| BTK | Agammaglobulinemia X-linked, type 1 |
| BUB1B | Mosaic variegated aneuploidy syndrome 1 |
| C10orf2 | Mitochondrial DNA depletion syndrome 7 (hepatocerebral type) |
| C12orf57 | Temtamy syndrome |
| C12orf65 | Combined oxidative phosphorylation deficiency, type 7; Spastic paraplegia, type 55, autosomal recessive |
| C19orf12 | Neurodegeneration with brain iron accumulation, type 4 |
| C1QA | C1q deficiency |
| C1QB | C1q deficiency |
| C1QBP | Combined oxidative phosphorylation deficiency 33 |
| C1QC | C1q deficiency |
| C1S | C1s deficiency |
| C2 | C2 deficiency |
| C2CD3 | Orofaciodigital syndrome, type 14 |
| C3 | Complement component 3 deficiency |
| C5 | Complement component 5 deficiency |
| C6 | Complement component 6 deficiency |
| C7 | Complement component 7 deficiency |
| C8B | Complement component 8 deficiency, type 2 |
| C8orf37 | Bardet-Biedl syndrome, type21 |
| CA12 | Hyperchlorhidrosis, isolated |
| CA2 | Osteopetrosis with renal tubular acidosis (osteopetrosis, autosomal recessive, type 3) |
| CA5A | Hyperammonemia due to carbonic anhydrase VA deficiency |
| CA8 | Cerebellar ataxia, impaired intellectual development and dysequilibrium syndrome 3 |
| CABP2 | Deafness, autosomal recessive, type 93 |
| CABP4 | Congenital stationary night blindness, type 2B |
| CACNA1D | Sinoatrial node dysfunction and deafness |
| CACNA2D4 | Retinal cone dystrophy 4 |
| CAD | Epileptic encephalopathy, early infantile, 50 |
| CALCRL | Lymphatic malformation 8 |
| CANT1 | Desbuquois dysplasia, type 1; Epiphyseal dysplasia, multiple, type 7 |
| CAPN1 | Spastic paraplegia, type 76 autosomal recessive |
| CAPN3 | Limb-girdle muscular dystrophy, type 1 (LGMD R1) |
| CARD11 | Immunodeficiency, type 11A |
| CARD9 | Candidiasis, familial, type 2 autosomal recessive |
| CARS2 | Combined oxidative phosphorylation deficiency 27 |
| CASK | FG syndrome 4 |
| CASP14 | Ichthyosis, congenital, autosomal recessive 12 |
| CASQ2 | Ventricular tachycardia, catecholaminergic polymorphic, type 2 |
| CASR | Hyperparathyroidism, neonatal |
| CAST | Peeling skin with leukonychia, acral punctate keratoses, cheilitis, and knuckle pads |
| CAT | Acatalasemia |
| CATSPER1 | Spermatogenic failure, type 7 |
| CAVIN1 | Lipodystrophy, congenital generalized, type 4 |
| CBLIF | Intrinsic factor deficiency |
| CBS | Homocystinuria due to cystathionine beta-synthase |
| CC2D1A | Intellectual developmental disorder, autosomal recessive 3 |
| CC2D2A | Joubert syndrome, type 9; Meckel syndrome, type 6; COACH syndrome, 2 |
| CCBE1 | Hennekam lymphangiectasia-lymphedema syndrome, type 1 |
| CCDC103 | Ciliary dyskinesia, primary, type 17 |
| CCDC115 | Congenital disorder of glycosylation, type IIo |
| CCDC174 | Hypotonia, infantile, with psychomotor retardation |
| CCDC39 | Ciliary dyskinesia, primary, type 14 |
| CCDC40 | Ciliary dyskinesia, primary, type 15 |
| CCDC65 | Ciliary dyskinesia, primary, type 27 |
| CCDC8 | 3M syndrome 3 |
| CCDC88C | Hydrocephalus, congenital, type 1 |
| CCN6 | Arthropathy, progressive pseudorheumatoid, of childhood |
| CCNO | Ciliary dyskinesia, primary, type 29 |
| CD19 | Immunodeficiency, common variable, type 3 |
| CD247 | Immunodeficiency, type 25 |
| CD27 | Lymphoproliferative syndrome 2 |
| CD2AP | Glomerulosclerosis, focal segmental, type 3 susceptibility to |
| CD320 | Methylmalonic aciduria, transient, due to transcobalamin receptor defect |
| CD36 | Platelet glycoprotein 4 deficiency |
| CD3D | Immunodeficiency, type 19 |
| CD3E | Immunodeficiency, type 18 |
| CD3G | Immunodeficiency, type 17 CD3 gamma deficient |
| CD40 | Immunodeficiency with hyper-IgM, type 3 |
| CD40LG | Hyper-IgM syndrome, type 1 (immunodeficiency, X-linked, with hyper-IgM, type 1) |
| CD55 | Complement hyperactivation, angiopathic thrombosis, and protein-losing enteropathy (CHAPLE) |
| CD59 | CD59 deficiency |
| CD79A | Agammaglobulinemia 3 |
| CD79B | Agammaglobulinemia 6 |
| CD81 | Immunodeficiency, common variable, type 6 |
| CD8A | CD8 deficiency, familial |
| CDAN1 | Dyserythropoietic anemia, congenital, type 1A |
| CDC14A | Deafness, autosomal recessive, type 105 |
| CDC45 | Meier-Gorlin syndrome 7 |
| CDCA7 | Immunodeficiency-centromeric instability-facial anomalies syndrome 3 |
| CDH11 | Elsahy-Waters syndrome |
| CDH23 | Deafness, autosomal recessive, type 12; Usher syndrome, type 1D |
| CDH3 | Ectodermal dysplasia, ectrodactyly, and macular dystrophy |
| CDHR1 | Cone-rod dystrophy, type 15 |
| CDIN1 | Dyserythropoietic anemia, congenital, type Ib |
| CDK10 | Al Kaissi syndrome |
| CDK5RAP2 | Primary microcephaly type 3 autosomal recessive |
| CDKL5 | Developmental and epileptic encephalopathy 2 |
| CDSN | Peeling skin syndrome 1 |
| CDT1 | Meier-Gorlin syndrome, type 4 |
| CEBPE | Specific granule deficiency |
| CENPF | Stromme syndrome |
| CENPJ | Primary microcephaly type 6 autosomal recessive |
| CEP104 | Joubert syndrome 25 |
| CEP120 | Short-rib thoracic dysplasia 13 with or without polydactyly |
| CEP135 | Microcephaly 8 primary, autosomal recessive |
| CEP152 | Primary microcephaly type 9 autosomal recessive |
| CEP164 | Nephronophthisis 15 |
| CEP19 | Morbid obesity and spermatogenic failure |
| CEP290 | Meckel syndrome, type 4; Joubert syndrome, type 5; Leber congenital amaurosis, type 10 |
| CEP41 | Joubert syndrome, type 15 |
| CEP55 | Multinucleated neurons, anhydramnios, renal dysplasia, cerebellar hypoplasia, and hydranencephaly |
| CEP57 | Mosaic variegated aneuploidy syndrome 2 |
| CEP78 | Cone-rod dystrophy and hearing loss |
| CEP83 | Nephronophthisis 18 |
| CERKL | Retinitis pigmentosa, type 26 |
| CERS3 | Ichthyosis, congenital, autosomal recessive 9 |
| CFAP43 | Spermatogenic failure, type 19 |
| CFAP53 | Heterotaxy, visceral, 6 autosomal recessive |
| CFD | Complement factor D deficiency |
| CFH | Complement factor H deficiency |
| CFI | Complement factor I deficiency |
| CFL2 | Nemaline myopathy, type 7 autosomal recessive |
| CFP | Properdin deficiency, X-linked |
| CFTR | Cystic fibrosis |
| CHAT | Myasthenic syndrome, congenital, type 6 presynaptic |
| CHKB | Muscular dystrophy, congenital, megaconial type |
| CHM | Choroideremia |
| CHMP1A | Pontocerebellar hypoplasia, type 8 |
| CHRNA1 | Multiple pterygium syndrome, lethal type |
| CHRNB1 | Myasthenic syndrome, congenital, 2C, associated with acetylcholine receptor deficiency |
| CHRND | Myasthenic syndrome, congenital, type 3B, fast-channel; Multiple pterygium syndrome, lethal type |
| CHRNE | Myasthenic syndrome, congenital, type 4B, fast-channel; Myasthenic syndrome, congenital, type 4C, associated with acetylcholine receptor deficiency |
| CHRNG | Multiple pterygium syndrome (MPS), Escobar type; MPS, lethal type |
| CHST14 | Ehlers-Danlos syndrome, musculocontractural, type 1 |
| CHST3 | Spondyloepiphyseal dysplasia with congenital joint dislocations |
| CHST6 | Macular corneal dystrophy |
| CHSY1 | Temtamy preaxial brachydactyly syndrome |
| CHUK | Cocoon syndrome |
| CIB2 | Deafness, autosomal recessive, type 48; Usher syndrome, type 1J |
| CIITA | Bare lymphocyte syndrome, type 2 complementation group A |
| CILK1 | Endocrine-cerebroosteodysplasia |
| CISD2 | Wolfram syndrome 2 |
| CIT | Microcephaly 17 primary, autosomal recessive |
| CKAP2L | Filippi syndrome |
| CLCF1 | Cold-induced sweating syndrome 2 |
| CLCN1 | Myotonia congenita, recessive |
| CLCN2 | Leukoencephalopathy with ataxia |
| CLCN5 | Dent disease 1 |
| CLCN7 | Osteopetrosis, autosomal recessive type 4 |
| CLCNKA | Bartter syndrome, type 4B, digenic |
| CLCNKB | Bartter syndrome, type 3; Bartter syndrome, type 4B, digenic |
| CLDN1 | Ichthyosis, leukocyte vacuoles, alopecia, and sclerosing cholangitis |
| CLDN10 | HELIX syndrome |
| CLDN14 | Deafness type 29 autosomal recessive |
| CLDN16 | Hypomagnesemia, type 3 renal |
| CLDN19 | Rena hypomagnesemia type 5 with ocular involvement |
| CLMP | Congenital short bowel syndrome |
| CLN3 | Ceroid lipofuscinosis, neuronal, type 3 |
| CLN5 | Ceroid lipofuscinosis, neuronal, type 5 |
| CLN6 | Ceroid lipofuscinosis, neuronal, type 6 |
| CLN8 | Ceroid lipofuscinosis, neuronal, type 8 |
| CLP1 | Pontocerebellar hypoplasia, type 10 |
| CLPB | 3-methylglutaconic aciduria, type 7 with cataracts, neurologic involvement and neutropenia |
| CLPP | Perrault syndrome 3 |
| CLRN1 | Usher syndrome, type 3A |
| CNGA1 | Retinitis pigmentosa type 49 |
| CNGA3 | Achromatopsia, type 2 |
| CNGB1 | Retinitis pigmentosa type 45 |
| CNGB3 | Achromatopsia, type 3 |
| CNNM2 | Hypomagnesemia, seizures, and intellectual developmental disorder |
| CNNM4 | Jalili syndrome |
| CNPY3 | Epileptic encephalopathy, early infantile, type 60 |
| CNTNAP1 | Lethal congenital contracture syndrome 7 |
| CNTNAP2 | Pitt-Hopkins like syndrome 1 |
| COA6 | Cardioencephalomyopathy, fatal infantile, due to cytochrome c oxidase deficiency 4 |
| COA8 | Mitochondrial complex IV deficiency, nuclear type 17 |
| COASY | Neurodegeneration with brain iron accumulation 6 |
| COG1 | Congenital disorder of glycosylation, type IIg |
| COG4 | Congenital disorder of glycosylation, type 2J |
| COG5 | Congenital disorder of glycosylation, type 2I |
| COG6 | Congenital disorder of glycosylation, type 2L; Shaheen syndrome |
| COG7 | Congenital disorder of glycosylation, type 2E |
| COG8 | Congenital disorder of glycosylation, type 2H |
| COL11A1 | Fibrochondrogenesis type 1 |
| COL11A2 | Otospondylomegaepiphyseal dysplasia, autosomal recessive |
| COL13A1 | Myasthenic syndrome, congenital, 19 |
| COL17A1 | Epidermolysis bullosa, junctional, non-Herlitz type |
| COL18A1 | Knobloch syndrome, type 1 |
| COL25A1 | Fibrosis of extraocular muscles, congenital, type 5 |
| COL27A1 | Steel syndrome |
| COL4A3 | Alport syndrome, autosomal recessive, type 2 |
| COL4A4 | Alport syndrome, autosomal recessive, type 2 |
| COL4A5 | Alport syndrome, X-linked |
| COL6A1 | Ullrich congenital muscular dystrophy, type 1 (Limb-girdle muscular dystrophy, type 22 [LGMD R22]) |
| COL6A2 | Ullrich congenital muscular dystrophy, type 1 (Limb-girdle muscular dystrophy, type 22 [LGMD R22]) |
| COL6A3 | Ullrich congenital muscular dystrophy, type 1 (Limb-girdle muscular dystrophy, type 22 [LGMD R22]) |
| COL7A1 | Dystrophic epidermolysis bullosa (DEB), Hallopeau-Siemens (HS) type and non-HS type; DEB pruriginosa; DEB pretibial |
| COL9A1 | Stickler syndrome, type 4 |
| COL9A2 | Stickler syndrome, type V |
| COLEC10 | 3MC syndrome 3 |
| COLEC11 | 3MC syndrome 2 |
| COLQ | Myasthenic syndrome, congenital, type 5 |
| COQ2 | Primary coenzyme Q10 deficiency, type 1 |
| COQ4 | Coenzyme Q10 deficiency, primary, type 7 |
| COQ6 | Coenzyme Q10 deficiency, primary, type 6 |
| COQ8A | Primary coenzyme Q10 deficiency, type 4 |
| COQ8B | Nephrotic syndrome, type 9 |
| COQ9 | Coenzyme Q10 deficiency, primary, type 5 |
| CORO1A | Immunodeficiency, type 8 |
| COX10 | Mitochondrial complex IV deficiency, nuclear type 3 |
| COX15 | Cardioencephalomyopathy, fatal infantile, due to cytochrome c oxidase deficiency, type 2; Leigh syndrome due to cytochrome c oxidase deficiency |
| COX20 | Mitochondrial complex IV deficiency, nuclear type 11 |
| COX6B1 | Mitochondrial complex IV deficiency, nuclear type 7 |
| CP | Aceruloplasminemia |
| CPA6 | Febrile seizures, familial, type 11 |
| CPAMD8 | Anterior segment dysgenesis, type 8 |
| CPLANE1 | Joubert syndrome 17 |
| CPLX1 | Epileptic encephalopathy, early infantile, 63 |
| CPS1 | Carbamoylphosphate synthetase 1 deficiency |
| CPT1A | Carnitine palmitoyltransferase type 1A deficiency, hepatic |
| CPT2 | Carnitine palmitoyltransferase type 2 deficiency, lethal neonatal; Carnitine palmitoyltransferase type 2 deficiency, infantile |
| CR2 | Immunodeficiency, common variable, type 7 |
| CRADD | intellectual developmental disorder, autosomal recessive, type 34 with variant lissencephaly |
| CRB1 | Retinitis pigmentosa, type 12; Leber congenital amaurosis, type 8 |
| CRB2 | Ventriculomegaly with cystic kidney disease |
| CRBN | intellectual developmental disorder, autosomal recessive, type 2 |
| CRIPT | Short stature with microcephaly and distinctive facies |
| CRLF1 | Cold-induced sweating syndrome type 1 |
| CRPPA | Muscular dystrophy-dystroglycanopathy, type A7; Muscular dystrophy-dystroglycanopathy, type C7 |
| CRTAP | Osteogenesis imperfecta, type 7 |
| CRYAA | Cataract 9 multiple types |
| CRYAB | Myopathy, myofibrillar, fatal infantile hypertonic, alpha-B crystallin-related; Cataract 16 multiple types |
| CRYBB1 | Cataract 17 |
| CRYBB3 | Cataract 22 |
| CSF2RB | Surfactant metabolism dysfunction, pulmonary, type 5 |
| CSF3R | Neutropenia, severe congenital, type 7 autosomal recessive |
| CSPP1 | Joubert syndrome 21 |
| CSTA | Peeling skin syndrome, type 4 |
| CSTB | Epilepsy, progressive myoclonic type 1A (Unverricht and Lundborg) |
| CTC1 | Cerebroretinal microangiopathy with calcifications and cysts |
| CTH | Cystathioninuria |
| CTNS | Nephropathic cystinosis |
| CTPS1 | Immunodeficiency, type 24 |
| CTSA | Galactosialidosis |
| CTSC | Haim-Munk syndrome; Papillon-Lefevre syndrome |
| CTSD | Ceroid lipofuscinosis, neuronal, type 10 |
| CTSF | Ceroid lipofuscinosis, neuronal, type 13 (Kufs type) |
| CTSK | Pycnodysostosis |
| CUBN | Megaloblastic anemia 1 (Imerslund-Grasbeck syndrome) |
| CUL4B | Intellectual developmental disorder, X-linked syndromic, Cabezas type |
| CUL7 | 3M syndrome 1 |
| CWC27 | Retinitis pigmentosa with or without skeletal anomalies |
| CWF19L1 | Spinocerebellar ataxia, autosomal recessive, type 17 |
| CYB5A | 46,XY disorder of sex development due to isolated 17,20-lyase deficiency |
| CYB5R3 | Methemoglobinemia, type 1; Methemoglobinemia, type 2 |
| CYBA | Chronic granulomatous disease, type 4 |
| CYBB | Chronic granulomatous disease, X-linked |
| CYC1 | Mitochondrial complex III deficiency, nuclear type 6 |
| CYP11A1 | 46,XY disorder of sex development-adrenal insufficiency due to CYP11A1 deficiency |
| CYP11B1 | Adrenal hyperplasia, congenital, due to 11-beta-hydroxylase deficiency |
| CYP11B2 | Hypoaldosteronism, congenital, due to CMO I deficiency |
| CYP17A1 | 17 alpha-hydroxylase/17,20-lyase deficiency |
| CYP19A1 | Aromatase deficiency |
| CYP1B1 | Glaucoma, primary congenital, type 3A |
| CYP21A2 | Congenital adrenal hyperplasia due to 21-hydroxylase deficiency |
| CYP24A1 | Hypercalcemia, infantile, type 1 |
| CYP26B1 | Craniosynostosis with radiohumeral fusions and other skeletal and craniofacial anomalies |
| CYP26C1 | Focal facial dermal dysplasia 4 |
| CYP27A1 | Cerebrotendinous xanthomatosis |
| CYP27B1 | Vitamin D-dependent rickets, type 1 |
| CYP2R1 | Rickets due to defect in vitamin D 25-hydroxylation |
| CYP2U1 | Spastic paraplegia, type 56 autosomal recessive |
| CYP4F22 | Ichthyosis, congenital, autosomal recessive, type 5 |
| CYP4V2 | Bietti crystalline corneoretinal dystrophy |
| CYP7B1 | Spastic paraplegia, type 5A, autosomal recessive |
| D2HGDH | D-2-hydroxyglutaric aciduria |
| DAG1 | Muscular dystrophy-dystroglycanopathy type A9; Muscular dystrophy-dystroglycanopathy type C9 |
| DARS1 | Hypomyelination with brainstem and spinal cord involvement and leg spasticity |
| DARS2 | Leukoencephalopathy with brain stem and spinal cord involvement and lactate elevation |
| DBH | Dopamine beta-hydroxylase deficiency |
| DBT | Maple syrup urine disease, type 2 |
| DCAF17 | Woodhouse-Sakati syndrome |
| DCC | Gaze palsy, familial horizontal, with progressive scoliosis, type 2 |
| DCDC2 | Sclerosing cholangitis, neonatal; Nephronophthisis 19 |
| DCHS1 | Van Maldergem syndrome 1 |
| DCLRE1C | Omenn syndrome; Severe combined immunodeficiency, Athabascan type |
| DCPS | Al-Raqad syndrome |
| DCX | Lissencephaly, X-linked, type 1 |
| DDB2 | Xeroderma pigmentosum, complementation group E |
| DDC | Aromatic L-amino acid decarboxylase deficiency |
| DDHD1 | Spastic paraplegia, type 28 autosomal recessive |
| DDHD2 | Spastic paraplegia, type 54 autosomal recessive |
| DDR2 | Spondylometaepiphyseal dysplasia, short limb-hand type |
| DDRGK1 | Spondyloepimetaphyseal dysplasia, Shohat type |
| DDX11 | Warsaw breakage syndrome |
| DDX59 | Orofaciodigital syndrome V |
| DENND5A | Epileptic encephalopathy, early infantile, 49 |
| DES | Myopathy, myofibrillar, type 1 |
| DGAT1 | Diarrhea 7 protein-losing enteropathy type |
| DGKE | Nephrotic syndrome, type 7 |
| DGUOK | DGUOK-related mitochondrial DNA depletion syndrome |
| DHCR24 | Desmosterolosis |
| DHCR7 | Smith-Lemli-Opitz syndrome |
| DHDDS | Retinitis pigmentosa, type 59 |
| DHFR | Megaloblastic anemia due to dihydrofolate reductase deficiency |
| DHH | 46,XY complete gonadal dysgenesis |
| DHODH | Miller syndrome |
| DHPS | Neurodevelopmental disorder with seizures and speech and walking impairment |
| DHTKD1 | 2-aminoadipic 2-oxoadipic aciduria |
| DIAPH1 | Seizures, cortical blindness, microcephaly syndrome |
| DIS3L2 | Perlman syndrome |
| DKC1 | Dyskeratosis congenita, X-linked |
| DLAT | Pyruvate dehydrogenase E2 deficiency |
| DLD | Dihydrolipoamide dehydrogenase deficiency |
| DLG3 | Intellectual developmental disorder, X-linked 90 |
| DLL3 | Spondylocostal dysostosis type 1 |
| DMD | Duchenne/Becker muscular dystrophy |
| DMGDH | Dimethylglycine dehydrogenase deficiency |
| DMP1 | Hypophosphatemic rickets, autosomal recessive |
| DMXL2 | Developmental and epileptic encephalopathy, type 81 |
| DNAAF1 | Ciliary dyskinesia, primary, type 13 |
| DNAAF2 | Ciliary dyskinesia, primary, type 10 |
| DNAAF3 | Ciliary dyskinesia, primary, type 2 |
| DNAAF4 | Ciliary dyskinesia, primary, type 25 |
| DNAAF5 | Ciliary dyskinesia, primary, type 18 |
| DNAH1 | Spermatogenic failure, type 18 |
| DNAH11 | Ciliary dyskinesia, primary, type 7 with or without situs inversus |
| DNAH5 | Ciliary dyskinesia, primary, type 3 with or without situs inversus |
| DNAH9 | Ciliary dyskinesia, primary, type 40 |
| DNAI1 | Ciliary dyskinesia, primary, type 1 with or without situs inversus |
| DNAI2 | Ciliary dyskinesia, primary, type 9 with or without situs inversus |
| DNAJB13 | Ciliary dyskinesia, primary, type 34 |
| DNAJB2 | Spinal muscular atrophy, distal, autosomal recessive, type 5 |
| DNAJC12 | Hyperphenylalaninemia, mild, non-BH4-deficient |
| DNAJC19 | 3-methylglutaconic aciduria, type 5 |
| DNAJC21 | Bone marrow failure syndrome, type 3 |
| DNAJC6 | Parkinson disease, type 19A, juvenile-onset; Parkinson disease, type 19B, early-onset |
| DNAL1 | Ciliary dyskinesia, primary, type 16 |
| DNASE1L3 | Systemic lupus erythematosus 16 |
| DNM1L | Encephalopathy due to defective mitochondrial and peroxisomal fission, type 1 |
| DNM2 | Lethal congenital contracture syndrome, type 5 |
| DNMT3B | Immunodeficiency-centromeric instability-facial anomalies syndrome, type 1 |
| DOCK2 | Immunodeficiency, type 40 |
| DOCK6 | Adams-Oliver syndrome 2 |
| DOCK7 | Epileptic encephalopathy, early infantile, 23 |
| DOCK8 | Hyper-IgE recurrent infection syndrome, autosomal recessive |
| DOK7 | Fetal akinesia deformation sequence, type 3; Myasthenic syndrome, congenital, type 10 |
| DOLK | Congenital disorder of glycosylation, type 1M |
| DONSON | Microcephaly, short stature, and limb abnormalities |
| DPAGT1 | Congenital disorder of glycosylation, type 1J; Myasthenic syndrome, congenital, type 13 |
| DPH1 | Developmental delay with short stature, dysmorphic features, and sparse hair |
| DPM1 | Congenital disorder of glycosylation, type 1E |
| DPM2 | Congenital disorder of glycosylation, type Iu |
| DPM3 | Congenital disorder of glycosylation, type Io |
| DPY19L2 | Spermatogenic failure, type 9 |
| DPYD | Dihydropyrimidine dehydrogenase deficiency |
| DPYS | Dihydropyrimidinuria |
| DRAM2 | Cone-rod dystrophy 21 |
| DRC1 | Ciliary dyskinesia, primary, type 21 |
| DSG1 | Erythroderma, congenital, with palmoplantar keratoderma, hypotrichosis, and hyper IgE |
| DSG4 | Hypotrichosis, type 6 |
| DSP | Cardiomyopathy, dilated, with woolly hair and keratoderma; Epidermolysis bullosa, lethal acantholytic |
| DST | Epidermolysis bullosa simplex, autosomal recessive, type 2 |
| DSTYK | Spastic paraplegia, type 23 autosomal recessive |
| DTNBP1 | Hermansky-Pudlak syndrome, type 7 |
| DUOX2 | Thyroid dyshormonogenesis, type 6 |
| DUOXA2 | Thyroid dyshormonogenesis, type 5 |
| DYM | Smith-McCort dysplasia; Dyggve-Melchior-Clausen disease |
| DYNC2H1 | Short-rib thoracic dysplasia, type 3 with or without polydactyly |
| DYNC2I1 | Short-rib thoracic dysplasia 8 with or without polydactyly |
| DYNC2I2 | Short-rib thoracic dysplasia 11 with or without polydactyly |
| DYNC2LI1 | Short-rib thoracic dysplasia 15 with polydactyly |
| DYNLT2B | Short-rib thoracic dysplasia 17 with or without polydactyly |
| DYSF | Miyoshi muscular dystrophy, type 1; Limb-girdle muscular dystrophy, type 2 (LGMD R2) |
| DZIP1L | Polycystic kidney disease 5 |
| EARS2 | Combined oxidative phosphorylation deficiency 12 |
| ECEL1 | Arthrogryposis, distal, type 5D |
| ECHS1 | Mitochondrial short-chain enoyl-CoA hydratase 1 deficiency |
| ECM1 | Urbach-Wiethe disease |
| EDA | Ectodermal dysplasia, type 1 hypohidrotic, X-linked |
| EDAR | Ectodermal dysplasia 10B, hypohidrotic/hair/tooth type |
| EDARADD | Ectodermal dysplasia 11B, hypohidrotic/hair/tooth type |
| EDN1 | Auriculocondylar syndrome, type 3 |
| EDN3 | Waardenburg syndrome, type 4B |
| EDNRB | ABCD syndrome |
| EFEMP2 | Cutis laxa, autosomal recessive, type 1B |
| EFL1 | Shwachman-Diamond syndrome 2 |
| EGFR | Inflammatory skin and bowel disease, neonatal, 2 |
| EGR2 | Dejerine-Sottas disease |
| EIF2AK3 | Wolcott-Rallison syndrome |
| EIF2AK4 | Pulmonary venoocclusive disease 2 |
| EIF2B1 | Leukoencephalopathy with vanishing white matter (VWM) |
| EIF2B2 | Leukoencephalopathy with vanishing white matter (VWM) |
| EIF2B3 | Leukoencephalopathy with vanishing white matter (VWM) |
| EIF2B4 | Leukoencephalopathy with vanishing white matter (VWM) |
| EIF2B5 | Leukoencephalopathy with vanishing white matter (VWM) |
| EIF4A3 | Robin sequence with cleft mandible and limb anomalies |
| ELAC2 | Combined oxidative phosphorylation deficiency 17 |
| ELMO2 | Vascular malformation, primary intraosseous |
| ELOVL4 | Ichthyosis, spastic quadriplegia, and intellectual developmental disorder |
| ELP2 | intellectual developmental disorder, autosomal recessive, type 58 |
| EMC1 | Cerebellar atrophy, visual impairment, and psychomotor retardation |
| EMD | Emery-Dreifuss muscular dystrophy, type 1 X-linked |
| EML1 | Band heterotopia |
| EMP2 | Nephrotic syndrome, type 10 |
| ENAM | Amelogenesis imperfecta, type 1C |
| ENO3 | Glycogen storage disease XIII |
| ENPP1 | Arterial calcification, generalized, of infancy, type 1 |
| ENTPD1 | Spastic paraplegia, type 64 autosomal recessive |
| EOGT | Adams-Oliver syndrome 4 |
| EPB41 | Elliptocytosis, type 1 |
| EPB42 | Spherocytosis, type 5 |
| EPCAM | Diarrhea 5 with tufting enteropathy, congenital |
| EPG5 | Vici syndrome |
| EPM2A | Epilepsy, progressive myoclonic, type 2A (Lafora) |
| EPRS1 | Leukodystrophy, hypomyelinating, type 15 |
| EPS8L2 | Deafness autosomal recessive, type 106 |
| ERAL1 | Perrault syndrome 6 |
| ERBB3 | Lethal congenital contractural syndrome, type 2 |
| ERCC1 | Cerebrooculofacioskeletal syndrome, type 4 |
| ERCC2 | Trichothiodystrophy, type 1 |
| ERCC3 | Trichothiodystrophy, type 2 |
| ERCC4 | Fanconi anemia, complementation group Q |
| ERCC5 | Cerebrooculofacioskeletal syndrome, type 3 |
| ERCC6 | Cockayne syndrome, type B; Cerebrooculofacioskeletal syndrome, type 1 |
| ERCC6L2 | Bone marrow failure syndrome, type 2 |
| ERCC8 | Cockayne syndrome, type A |
| ERLIN1 | Spastic paraplegia, type 62 autosomal recessive |
| ERLIN2 | Spastic paraplegia, type 18 autosomal recessive |
| ESCO2 | Roberts syndrome; Juberg-Hayward syndrome |
| ESPN | Deafness, autosomal recessive, type 36 |
| ESR1 | Estrogen resistance |
| ESRRB | Deafness, autosomal recessive, type 35 |
| ETFA | Glutaric acidemia, type 2A |
| ETFB | Glutaric acidemia, type 2B |
| ETFDH | Glutaric acidemia, type 2C |
| ETHE1 | Ethylmalonic encephalopathy |
| EVC | Ellis-van Creveld syndrome |
| EVC2 | Ellis-van Creveld syndrome |
| EXOSC3 | Pontocerebellar hypoplasia, type 1B |
| EXPH5 | Epidermolysis bullosa, nonspecific, autosomal recessive |
| EXT1 | Chondrosarcoma |
| EXTL3 | Immunoskeletal dysplasia with neurodevelopmental abnormalities |
| EYS | Retinitis pigmentosa, type 25 |
| F10 | Factor X deficiency |
| F11 | Factor XI deficiency |
| F13A1 | Factor XIIIA deficiency |
| F13B | Factor XIIIB deficiency |
| F2 | Prothrombin deficiency |
| F5 | Factor V deficiency |
| F7 | Factor VII deficiency |
| F8 | Hemophilia A |
| F9 | Hemophilia B |
| FA2H | Spastic paraplegia, type 35 autosomal recessive |
| FADD | Infections, recurrent, with encephalopathy, hepatic dysfunction, and cardiovascular malformations |
| FAH | Tyrosinemia, type 1 |
| FAM126A | Leukodystrophy, hypomyelinating, type 5 |
| FAM161A | Retinitis pigmentosa, type 28 |
| FAM20A | Amelogenesis imperfecta, type 1G (Enamel-renal syndrome) |
| FAM20C | Raine syndrome |
| FAN1 | Interstitial nephritis, karyomegalic |
| FANCA | Fanconi anemia, complementation group A |
| FANCB | Fanconi anemia, complementation group B |
| FANCC | Fanconi anemia, complementation group C |
| FANCD2 | Fanconi anemia, complementation group D2 |
| FANCE | Fanconi anemia, complementation group E |
| FANCF | Fanconi anemia, complementation group F |
| FANCG | Fanconi anemia, complementation group G |
| FANCI | Fanconi anemia, complementation group I |
| FANCL | Fanconi anemia, complementation group L |
| FANCM | Spermatogenic failure, type 28 |
| FAR1 | Peroxisomal fatty acyl-CoA reductase 1 disorder |
| FARS2 | Combined oxidative phosphorylation deficiency 14; Spastic paraplegia, type 77 autosomal recessive |
| FASTKD2 | Combined oxidative phosphorylation deficiency 44 |
| FAT4 | Hennekam lymphangiectasia-lymphedema syndrome 2 |
| FBLN5 | Cutis laxa, autosomal recessive, type 1A |
| FBP1 | Fructose-1,6-bisphosphatase deficiency |
| FBXL4 | Mitochondrial DNA depletion syndrome 13 (encephalomyopathic type) |
| FBXO7 | Parkinson disease, type 15 autosomal recessive |
| FDXR | Auditory neuropathy and optic atrophy |
| FECH | Protoporphyria, erythropoietic, autosomal recessive |
| FERMT1 | Kindler syndrome |
| FERMT3 | Leukocyte adhesion deficiency, type 3 |
| FEZF1 | Hypogonadotropic hypogonadism type 22 with or without anosmia |
| FGA | Afibrinogenemia, congenital |
| FGB | Congenital afibrinogenemia |
| FGD1 | Aarskog-Scott syndrome; intellectual developmental disorder, X-linked syndromic, type 16 |
| FGD4 | Charcot-Marie-Tooth disease, type 4H |
| FGF23 | Tumoral calcinosis, hyperphosphatemic, familial, type 2 |
| FGF3 | Deafness, congenital with inner ear agenesis, microtia, and microdontia |
| FGG | Afibrinogenemia, congenital; Hypofibrinogenemia, congenital |
| FH | Fumarase deficiency |
| FHL1 | Emery-Dreifuss muscular dystrophy 6, X-linked |
| FIBP | Thauvin-Robinet-Faivre syndrome |
| FIG4 | Charcot-Marie-Tooth disease, type 4J; Yunis-Varon syndrome |
| FKBP10 | Bruck syndrome 1 |
| FKBP14 | Ehlers-Danlos syndrome, kyphoscoliotic type, 2 |
| FKRP | Muscular dystrophy-dystroglycanopathy, type 5A (Walker-Warburg syndrome); Type 5B; Type 5C (limb-girdle muscular dystrophy, type 9 [LGMDR9]) |
| FKTN | Muscular dystrophy-dystroglycanopathy, type 4A (Walker-Warburg syndrome); Type 4B; Type 4C (limb-girdle muscular dystrophy, type 13 [LGMD R13]) |
| FLAD1 | Lipid storage myopathy due to flavin adenine dinucleotide synthetase deficiency |
| FLG | Ichthyosis vulgaris |
| FLI1 | Bleeding disorder, platelet-type, type 21 |
| FLNA | X-linked otopalatodigital (X-OPD) spectrum disorders; |
| FLNB | Spondylocarpotarsal synostosis syndrome |
| FLVCR1 | Posterior column ataxia-retinitis pigmentosa syndrome |
| FLVCR2 | Proliferative vasculopathy and hydranencephaly-hydrocephaly syndrome |
| FMN2 | intellectual developmental disorder, autosomal recessive, type 47 |
| FMO3 | Trimethylaminuria |
| FMR1 | Fragile X syndrome |
| FOLR1 | Neurodegeneration due to cerebral folate transport deficiency |
| FOXE1 | Bamforth-Lazarus syndrome |
| FOXE3 | Anterior segment dysgenesis, type 2 multiple subtypes |
| FOXN1 | T-cell immunodeficiency, congenital alopecia and nail dystrophy |
| FOXP3 | Immunodysregulation, polyendocrinopathy, and enteropathy, X-linked |
| FOXRED1 | Mitochondrial complex I deficiency, nuclear type 19 |
| FRAS1 | Fraser syndrome, type 1 |
| FREM1 | Manitoba oculotrichoanal syndrome |
| FREM2 | Fraser syndrome, type 2 |
| FRRS1L | Epileptic encephalopathy, early infantile, 37 |
| FSHB | Hypogonadotropic hypogonadism, type 24 without anosmia |
| FSHR | Ovarian dysgenesis 1 |
| FTCD | Glutamate formiminotransferase deficiency |
| FTL | L-ferritin deficiency |
| FTO | Growth retardation, developmental delay, facial dysmorphism |
| FTSJ1 | Intellectual developmental disorder, X-linked 9 |
| FUCA1 | Fucosidosis |
| FUT8 | Congenital disorder of glycosylation with defective fucosylation, type 1 |
| FXN | Friedreich ataxia |
| FYCO1 | Cataract 18 |
| FZD6 | Nail disorder, nonsyndromic congenital, type 10 (claw-shaped nails) |
| G6PC | Glycogen storage disease, type 1A |
| G6PC3 | Dursun syndrome |
| G6PD | Hemolytic anemia, G6PD deficient (favism) |
| GAA | Glycogen storage disease, type 2 |
| GALC | Krabbe disease |
| GALE | Galactose epimerase deficiency |
| GALK1 | Galactokinase deficiency with cataracts |
| GALNS | Mucopolysaccharidosis, type 4A |
| GALNT3 | Tumoral calcinosis, hyperphosphatemic, familial, type 1 |
| GALT | Galactosemia |
| GAMT | Cerebral creatine deficiency syndrome, type 2 |
| GAN | Giant axonal neuropathy, type 1 |
| GAS8 | Ciliary dyskinesia, primary, type 33 |
| GATM | Cerebral creatine deficiency syndrome, type 3 |
| GBA | Gaucher Disease |
| GBA2 | Spastic paraplegia, type 46 autosomal recessive |
| GBE1 | Glycogen storage disease, type 4 |
| GCDH | Glutaricaciduria, type 1 |
| GCH1 | Hyperphenylalaninemia, BH4-deficient, type B |
| GCK | Permanent neonatal diabetes mellitus (PNDM) |
| GCM2 | Hypoparathyroidism, familial isolated (FIH) 2 |
| GCNT2 | Cataract 13 with adult i phenotype |
| GCSH | ?Glycine encephalopathy |
| GDAP1 | Charcot-Marie-Tooth disease, recessive intermediate, type A |
| GDF1 | Right atrial isomerism (Ivemark syndrome) |
| GDF5 | Chondrodysplasia, Grebe type |
| GDF6 | Leber congenital amaurosis, type 17 |
| GDI1 | Intellectual developmental disorder, X-linked 41 |
| GFER | Myopathy, mitochondrial progressive, with congenital cataract, hearing loss, and developmental delay |
| GFM1 | Combined oxidative phosphorylation deficiency, type 1 |
| GFPT1 | Myasthenia, congenital, type 12 with tubular aggregates |
| GGCX | Vitamin K-dependent clotting factors, combined deficiency of, type 1 |
| GH1 | Growth hormone deficiency, isolated, type 1A; Kowarski syndrome |
| GHR | Laron dwarfism |
| GHRHR | Growth hormone deficiency, isolated, type 1B |
| GHSR | Growth hormone deficiency, isolated partial |
| GINS1 | Immunodeficiency, type 55 |
| GIPC3 | Deafness, autosomal recessive, type 15 |
| GJA1 | Craniometaphyseal dysplasia, autosomal recessive |
| GJB1 | Charcot-Marie-Tooth neuropathy, X-linked dominant, type 1 |
| GJB2 | Deafness, autosomal recessive, type 1A; Deafness, digenic, GJB2/GJB6 |
| GJB6 | Deafness, autosomal recessive, type 1B; Deafness, digenic GJB2/GJB6 |
| GJC2 | Spastic paraplegia, type 44 autosomal recessive |
| GK | Glycerol kinase deficiency |
| GLA | Fabry disease |
| GLB1 | GM1-gangliosidosis, types 1-3; Mucopolysaccharidosis, type 4B (Morquio) |
| GLDC | Glycine encephalopathy |
| GLDN | Lethal congenital contracture syndrome 11 |
| GLE1 | Lethal congenital contracture syndrome, type 1; Congenital arthrogryposis with anterior horn cell disease |
| GLIS2 | Nephronophthisis, type 7 |
| GLIS3 | Diabetes mellitus, neonatal, with congenital hypothyroidism |
| GLRA1 | Hyperekplexia, type 1 |
| GLRB | Hyperekplexia, type 2 |
| GLRX5 | Anemia, sideroblastic, type 3 pyridoxine-refractory; Spasticity, childhood-onset, with hyperglycinemia |
| GLUL | Glutamine deficiency, congenital |
| GLYCTK | D-glyceric aciduria |
| GM2A | GM2-gangliosidosis, AB variant |
| GMPPA | Alacrima, achalasia, and intellectual developmental disorder syndrome |
| GMPPB | Muscular dystrophy-dystroglycanopathy 14 |
| GNAT1 | Night blindness, congenital stationary, type 1G |
| GNAT2 | Achromatopsia, type 4 |
| GNB5 | Intellectual developmental disorder with cardiac arrhythmia; Language delay and ADHD/cognitive impairment with or without cardiac arrhythmia |
| GNE | Inclusion body myopathy, type 2 (Nonaka myopathy) |
| GNMT | Glycine N-methyltransferase deficiency |
| GNPAT | Rhizomelic chondrodysplasia punctata, type 2 |
| GNPTAB | Mucolipidosis 2 alpha/beta; Mucolipidosis 3 alpha/beta |
| GNPTG | Mucolipidosis III gamma |
| GNRHR | Hypogonadotropic hypogonadism, type 7 without anosmia |
| GNS | Mucopolysaccharidosis, type 3D (Sanfilippo syndrome D) |
| GORAB | Geroderma osteodysplasticum |
| GOSR2 | Epilepsy, progressive myoclonic, type 6 |
| GOT2 | Epileptic encephalopathy, early infantile, 82 |
| GP1BA | Bernard-Soulier syndrome, type A1 |
| GP1BB | Bernard-Soulier syndrome, type B |
| GP6 | Bleeding disorder, platelet-type, type 11 |
| GP9 | Bernard-Soulier syndrome, type C |
| GPAA1 | Glycosylphosphatidylinositol biosynthesis defect 15 |
| GPC3 | Simpson-Golabi-Behmel syndrome, type 1 |
| GPC6 | Omodysplasia, type 1 |
| GPD1 | Hypertriglyceridemia, transient infantile |
| GPHN | Molybdenum cofactor deficiency C |
| GPI | Hemolytic anemia, nonspherocytic, due to glucose phosphate isomerase deficiency |
| GPIHBP1 | Hyperlipoproteinemia, type 1D |
| GPR143 | Ocular albinism, type 1 (Nettleship-Falls type) |
| GPR179 | Night blindness, congenital stationary (complete), type 1E, autosomal recessive |
| GPR68 | Amelogenesis imperfecta, type 2A6 (hypomaturation type) |
| GPR98 | Usher syndrome, type 2C |
| GPSM2 | Chudley-McCullough syndrome |
| GPT2 | intellectual developmental disorder, autosomal recessive 49 |
| GPX4 | Spondylometaphyseal dysplasia, Sedaghatian type |
| GRHL2 | Ectodermal dysplasia/short stature syndrome |
| GRHPR | Hyperoxaluria, primary, type 2 |
| GRIA3 | Intellectual developmental disorder, X-linked syndromic, Wu type |
| GRID2 | Spinocerebellar ataxia, autosomal recessive, type 18 |
| GRIK2 | Intellectual developmental disorder, autosomal recessive 6 |
| GRIN1 | Neurodevelopmental disorder with or without hyperkinetic movements and seizures, autosomal recessive |
| GRIP1 | Fraser syndrome 3 |
| GRK1 | Oguchi disease-2 |
| GRM1 | Spinocerebellar ataxia, autosomal recessive, type 13 |
| GRM6 | Night blindness, congenital stationary (complete), type 1B, autosomal recessive |
| GRN | Ceroid lipofuscinosis, neuronal, type 11 |
| GRXCR1 | Deafness, autosomal recessive, type 25 |
| GSC | Short stature, auditory canal atresia, mandibular hypoplasia, skeletal abnormalities |
| GSS | Glutathione synthetase deficiency |
| GTF2H5 | Trichothiodystrophy, type 3 photosensitive |
| GTPBP2 | Jaberi-Elahi syndrome |
| GTPBP3 | Combined oxidative phosphorylation deficiency 23 |
| GUCY2C | Meconium ileus |
| GUCY2D | Leber congenital amaurosis, type 1 |
| GUF1 | Epileptic encephalopathy, early infantile, 40 |
| GUSB | Mucopolysaccharidosis, type 7 |
| GYG1 | Polyglucosan body myopathy, type 2 |
| GYS1 | Glycogen storage disease, type 0 muscle |
| GYS2 | Glycogen storage disease, type 0 liver |
| GZF1 | Joint laxity, short stature, and myopia |
| H6PD | Cortisone reductase deficiency 1 |
| HAAO | Vertebral, cardiac, renal, and limb defects syndrome 1 |
| HACE1 | Spastic paraplegia and psychomotor retardation with or without seizures |
| HADH | 3-hydroxyacyl-CoA dehydrogenase deficiency |
| HADHA | Long-chain 3-hydroxyl-CoA dehydrogenase (LCHAD) deficiency; Mitochondrial trifunctional protein deficiency |
| HADHB | Mitochondrial trifunctional protein deficiency |
| HAMP | Hemochromatosis, type 2B |
| HARS1 | Usher syndrome, type 3B |
| HAX1 | Neutropenia, severe congenital, type 3 autosomal recessive |
| HBA1 | Alpha-thalassemia |
| HBA2 | Alpha-thalassemia |
| HBB | Beta-thalassemia, Sickle cell anemia and other HBB-related hemoglobinopathies |
| HCCS | Linear skin defects with multiple congenital anomalies 1 |
| HCFC1 | intellectual developmental disorder, X-linked 3 (methylmalonic acidemia and homocysteinemia, cblX type ) |
| HELLS | Immunodeficiency-centromeric instability-facial anomalies syndrome 4 |
| HEPACAM | Megalencephalic leukoencephalopathy with subcortical cysts 2A |
| HERC1 | Macrocephaly, dysmorphic facies, and psychomotor retardation |
| HERC2 | intellectual developmental disorder, autosomal recessive, type 38 |
| HES7 | Spondylocostal dysostosis, type 4 autosomal recessive |
| HESX1 | Growth hormone deficiency with pituitary anomalies |
| HEXA | Tay-Sachs disease |
| HEXB | Sandhoff disease, infantile, juvenile, and adult forms |
| HFE | Hemochromatosis |
| HFE2 | Hemochromatosis, type 2A |
| HFM1 | Premature ovarian failure 9 |
| HGD | Alkaptonuria |
| HGF | Deafness, autosomal recessive, type 39 |
| HGSNAT | Mucopolysaccharidosis type 3C (Sanfilippo syndrome C) |
| HIBCH | 3-hydroxyisobutryl-CoA hydrolase deficiency |
| HIKESHI | Leukodystrophy, hypomyelinating, type 13 |
| HINT1 | Neuromyotonia and axonal neuropathy, autosomal recessive |
| HJV | Hemochromatosis, type 2A |
| HK1 | Charcot-Marie-Tooth disease, type 4G |
| HLCS | Holocarboxylase synthetase deficiency |
| HMGCL | HMG-CoA lyase deficiency |
| HMGCS2 | HMG-CoA synthase-2 deficiency |
| HMOX1 | Heme oxygenase-1 deficiency |
| HMX1 | Oculoauricular syndrome |
| HNMT | intellectual developmental disorder, autosomal recessive, type 51 |
| HOGA1 | Hyperoxaluria, primary, type 3 |
| HOXA1 | Athabaskan brainstem dysgenesis syndrome |
| HOXB1 | Facial paresis, hereditary congenital, 3 |
| HOXC13 | Ectodermal dysplasia 9 hair/nail type |
| HPCA | Dystonia 2 torsion, autosomal recessive |
| HPD | Tyrosinemia, type 3 |
| HPGD | Hypertrophic osteoarthropathy, primary, type 1 (pachydermoperiostosis) |
| HPRT1 | Lesch-Nyhan syndrome |
| HPS1 | Hermansky-Pudlak syndrome, type 1 |
| HPS3 | Hermansky-Pudlak syndrome, type 3 |
| HPS4 | Hermansky-Pudlak syndrome, type 4 |
| HPS5 | Hermansky-Pudlak syndrome, type 5 |
| HPS6 | Hermansky-Pudlak syndrome, type 6 |
| HPSE2 | Urofacial syndrome, type 1 |
| HR | Alopecia universalis; Atrichia with papular lesions |
| HSD11B2 | Apparent mineralocorticoid excess |
| HSD17B10 | HSD10 mitochondrial disease |
| HSD17B3 | 46,XY disorder of sex development due to 17-beta-hydroxysteroid dehydrogenase 3 deficiency |
| HSD17B4 | D-bifunctional protein deficiency |
| HSD3B2 | Adrenal hyperplasia, congenital, due to 3-beta-hydroxysteroid dehydrogenase 2 deficiency |
| HSD3B7 | Bile acid synthesis defect, congenital, type 1 |
| HSPA9 | Even-plus syndrome |
| HSPD1 | Leukodystrophy, hypomyelinating, type 4 |
| HSPG2 | Dyssegmental dysplasia, Silverman-Handmaker type |
| HTRA1 | CARASIL syndrome |
| HTRA2 | 3-methylglutaconic aciduria, type 8 |
| HUWE1 | Intellectual developmental disorder, X-linked syndromic, Turner type |
| HYAL1 | ?Mucopolysaccharidosis, type 9 |
| HYDIN | Ciliary dyskinesia, primary, type 5 |
| HYLS1 | Hydrolethalus syndrome |
| IARS1 | Growth retardation, intellectual developmental disorder, hypotonia, and hepatopathy |
| IBA57 | Multiple mitochondrial dysfunctions syndrome 3 |
| ICOS | Immunodeficiency, common variable, 1 |
| IDH3B | Retinitis pigmentosa, type 46 |
| IDS | Mucopolysaccharidosis, type 2 |
| IDUA | Mucopolysaccharidosis type 1 |
| IER3IP1 | Microcephaly, epilepsy, and diabetes syndrome |
| IFNGR1 | Immunodeficiency, type 27A, mycobacteriosis |
| IFNGR2 | Immunodeficiency, type 28 mycobacteriosis |
| IFT122 | Cranioectodermal dysplasia 1 |
| IFT140 | Retinitis pigmentosa, type 80; Short-rib thoracic dysplasia 9 with or without polydactyly |
| IFT172 | Short-rib thoracic dysplasia 10 with or without polydactyly |
| IFT43 | Short-rib thoracic dysplasia 18 with polydactyly |
| IFT52 | Short-rib thoracic dysplasia 16 with or without polydactyly |
| IFT80 | Short-rib thoracic dysplasia, type 2 with or without polydactyly |
| IFT81 | Short-rib thoracic dysplasia 19 with or without polydactyly |
| IGBP1 | Corpus callosum, agenesis of, with impaired intellectual development, ocular coloboma and micrognathia |
| IGF1 | Growth retardation with deafness and intellectual developmental disorder due to IGF1 deficiency |
| IGF1R | Insulin-like growth factor I, resistance to |
| IGFALS | Acid-labile subunit deficiency |
| IGFBP7 | Retinal arterial macroaneurysm with supravalvular pulmonic stenosis |
| IGHMBP2 | Charcot-Marie-Tooth disease, axonal, type 2S |
| IGLL1 | Agammaglobulinemia 2 |
| IGSF1 | Hypothyroidism, central, and testicular enlargement |
| IHH | Acrocapitofemoral dysplasia |
| IKBKAP | Dysautonomia, familial |
| IKBKB | Immunodeficiency, type 15 |
| IKBKG | Ectodermal dysplasia and immunodeficiency 1 |
| IL10RA | Inflammatory bowel disease, type 28 early onset, autosomal recessive |
| IL10RB | Inflammatory bowel disease, type 25 early onset, autosomal recessive |
| IL11RA | Craniosynostosis and dental anomalies |
| IL12B | Immunodeficiency, type 29 mycobacteriosis |
| IL12RB1 | Immunodeficiency, type 30 |
| IL17RA | Immunodeficiency, type 51 |
| IL17RC | Candidiasis, familial, 9 |
| IL1RAPL1 | Intellectual developmental disorder, X-linked 21 |
| IL1RN | Sterile multifocal osteomyelitis with periostitis and pustulosis |
| IL21R | Immunodeficiency, type 56 |
| IL2RA | Immunodeficiency, type 41 with lymphoproliferation and autoimmunity |
| IL2RG | Severe combined immunodeficiency, X-linked |
| IL36RN | Psoriasis, type 14 pustular |
| IL7R | Severe combined immunodeficiency, T-cell negative, B-cell/natural killer cell-positive type |
| ILDR1 | Deafness, autosomal recessive, type 42 |
| IMPA1 | intellectual developmental disorder, autosomal recessive 59 |
| IMPG2 | Retinitis pigmentosa, type 56 |
| INPP5E | Joubert syndrome, type 1 |
| INPP5K | Muscular dystrophy, congenital, with cataracts and intellectual disability |
| INPPL1 | Opsismodysplasia |
| INS | Permanent neonatal diabetes mellitus (PNDM) |
| INSR | Diabetes mellitus, insulin-resistant, with acanthosis nigricans, type A |
| INTS1 | Neurodevelopmental disorder with cataracts, poor growth, and dysmorphic facies |
| INVS | Nephronophthisis, type 2 infantile |
| IQCB1 | Senior-Loken syndrome, type 5 |
| IQCE | Polydactyly, postaxial, type A7 |
| IQSEC2 | Intellectual developmental disorder, X-linked 1 |
| IRAK4 | Immunodeficiency, type 67 (IRAK4 deficiency) |
| IRF8 | Immunodeficiency, type 32B, monocyte and dendritic cell deficiency |
| IRX5 | Hamamy syndrome |
| ISCA1 | Multiple mitochondrial dysfunctions syndrome 5 |
| ISCA2 | Multiple mitochondrial dysfunctions syndrome 4 |
| ISCU | Myopathy with lactic acidosis, hereditary |
| ISG15 | Immunodeficiency, type 38 |
| ITCH | Autoimmune disease, multisystem, with facial dysmorphism |
| ITGA2B | Glanzmann thrombasthenia |
| ITGA3 | Interstitial lung disease, nephrotic syndrome, and epidermolysis bullosa, congenital |
| ITGA6 | Epidermolysis bullosa, junctional, with pyloric stenosis |
| ITGA7 | Muscular dystrophy, congenital, due to ITGA7 deficiency |
| ITGA8 | Renal hypodysplasia/aplasia 1 |
| ITGB2 | Leukocyte adhesion deficiency |
| ITGB3 | Glanzmann thrombasthenia |
| ITGB4 | Epidermolysis bullosa, junctional, with pyloric atresia |
| ITGB6 | Amelogenesis imperfecta, type 1H |
| ITK | Lymphoproliferative syndrome 1 |
| ITPA | Epileptic encephalopathy, early infantile, type 35 |
| ITPR1 | Gillespie syndrome |
| IVD | Isovaleric acidemia |
| IYD | Thyroid dyshormonogenesis, type 4 |
| JAGN1 | Neutropenia, severe congenital, 6 autosomal recessive |
| JAK3 | Severe Combined Immunodeficiency, autosomal recessive, T-negative/B-positive type |
| JAM3 | Hemorrhagic destruction of the brain, subependymal calcification, and cataracts |
| JUP | Naxos disease |
| KANK2 | Nephrotic syndrome, type 16 |
| KARS1 | Deafness, autosomal recessive, type 89 |
| KATNB1 | Lissencephaly 6 with microcephaly |
| KATNIP | Joubert syndrome 26 |
| KCNE1 | Jervell and Lange-Nielsen syndrome 2 |
| KCNJ1 | Bartter syndrome, type 2 |
| KCNJ10 | SESAME syndrome |
| KCNJ11 | Hyperinsulinemic hypoglycemia, type 2 (congenital hyperinsulinism); Permanent neonatal diabetes mellitus (PNDM) |
| KCNJ13 | Leber congenital amaurosis, type 16 |
| KCNV2 | Retinal cone dystrophy, type 3B |
| KCTD7 | Epilepsy, progressive myoclonic, type 3 with or without intracellular inclusions |
| KDM5C | Intellectual developmental disorder, X-linked syndromic, Claes-Jensen type |
| KDSR | Erythrokeratodermia variabilis et progressiva 4 |
| KERA | Cornea plana 2 autosomal recessive |
| KHDC3L | Hydatidiform mole, recurrent, type 2 |
| KIAA0586 | Short-rib thoracic dysplasia 14 with polydactyly |
| KIAA0753 | Orofaciodigital syndrome, type 15 |
| KIAA1109 | Alkuraya-Kucinskas syndrome |
| KIAA1549 | Retinitis pigmentosa, type 86 |
| KIAA2022 | Intellectual developmental disorder, X-linked 98 |
| KIF14 | Microcephaly 20 primary, autosomal recessive; ?Meckel syndrome 12 |
| KIF1A | Neuropathy, hereditary sensory, type 2C; Spastic paraplegia, type 30 autosomal recessive |
| KIF1C | Spastic ataxia 2 autosomal recessive |
| KIF7 | Acrocallosal syndrome; Joubert syndrome, type 12 |
| KIFBP | Goldberg-Shprintzen megacolon syndrome |
| KISS1R | Hypogonadotropic hypogonadism, type 8 with or without anosmia |
| KIZ | Retinitis pigmentosa 69 |
| KLHL3 | Pseudohypoaldosteronism, type 2D |
| KLHL40 | Nemaline myopathy 8 autosomal recessive |
| KLHL41 | Nemaline myopathy 9 |
| KLHL7 | Cold-induced sweating syndrome 3 |
| KLK4 | Amelogenesis imperfecta, type 2A1 (hypomaturation type) |
| KLKB1 | Fletcher factor (prekallikrein) deficiency |
| KNL1 | Microcephaly 4 primary, autosomal recessive |
| KPTN | intellectual developmental disorder, autosomal recessive 41 |
| KREMEN1 | Ectodermal dysplasia 13 hair/tooth type |
| KRT10 | Epidermolytic hyperkeratosis |
| KRT14 | Epidermolysis bullosa simplex, autosomal recessive, type 1 |
| KRT25 | Woolly hair, autosomal recessive 3 |
| KRT5 | Epidermolysis bullosa simplex, autosomal recessive, type 1 |
| KRT85 | Ectodermal dysplasia 4 hair/nail type |
| KY | Myopathy, myofibrillar, type 7 |
| KYNU | Vertebral, cardiac, renal, and limb defects syndrome, type 2 |
| L1CAM | L1 Syndrome |
| L2HGDH | L-2-hydroxyglutaric aciduria |
| LAMA1 | Poretti-Boltshauser syndrome |
| LAMA2 | LAMA2-related muscular dystrophy |
| LAMA3 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LAMB1 | Lissencephaly, type 5 |
| LAMB2 | Pierson syndrome; Nephrotic syndrome, type 5 with or without ocular abnormalities |
| LAMB3 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LAMC2 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LAMC3 | Cortical malformations, occipital |
| LAMP2 | Danon disease |
| LARGE1 | Muscular dystrophy-dystroglycanopathy, type 6A and 6B |
| LARP7 | Alazami syndrome |
| LARS1 | Infantile liver failure syndrome 1 (ILFS1) |
| LARS2 | Perrault syndrome, type 4 |
| LAT | Immunodeficiency, type 52 |
| LBR | Greenberg skeletal dysplasia |
| LCA5 | Leber congenital amaurosis, type 5 |
| LCAT | Familial LCAT deficiency; Fish-eye disease |
| LCK | Immunodeficiency, type 22 |
| LCT | Lactase deficiency, congenital |
| LDHA | Glycogen storage disease type 11 |
| LDLRAP1 | Hypercholesterolemia, familial, autosomal recessive |
| LEMD2 | Cataract 46 juvenile-onset |
| LEP | Obesity, morbid, due to leptin deficiency |
| LEPR | Obesity, morbid, due to leptin receptor deficiency |
| LGI4 | Arthrogryposis multiplex congenita, neurogenic, with myelin defect |
| LHB | Hypogonadotropic hypogonadism, type 23 with or without anosmia |
| LHCGR | Leydig cell hypoplasia |
| LHFPL5 | Deafness, autosomal recessive, type 67 |
| LHX3 | Pituitary hormone deficiency, combined, type 3 |
| LIAS | Hyperglycinemia, lactic acidosis, and seizures |
| LIFR | Stuve-Wiedemann syndrome / Schwartz-Jampel type 2 syndrome |
| LIG4 | LIG4 syndrome |
| LIM2 | Cataract 19 multiple types |
| LINS1 | intellectual developmental disorder, autosomal recessive, type 27 |
| LIPA | Lysosomal Acid Lipase Deficiency |
| LIPE | Lipodystrophy, familial partial, type 6 |
| LIPH | Hypotrichosis, type 7 or woolly hair, autosomal recessive, type 2 with or without hypotrichosis |
| LIPN | Ichthyosis, congenital, autosomal recessive 8 |
| LIPT1 | Lipoyltransferase 1 deficiency |
| LIPT2 | Encephalopathy, neonatal severe, with lactic acidosis and brain abnormalities |
| LMAN1 | Combined deficiency of factor V and factor VIII, type 1 |
| LMBRD1 | Methylmalonic aciduria and homocystinuria, cblF type |
| LMF1 | Lipase deficiency, combined |
| LMNA | Cardiomyopathy, dilated, 1A |
| LMOD3 | Nemaline myopathy 10 |
| LONP1 | CODAS syndrome |
| LOXHD1 | Deafness, autosomal recessive, type 77 |
| LPAR6 | Hypotrichosis, type 8 or woolly hair, autosomal recessive, type 1 with or without hypotrichosis |
| LPIN1 | Myoglobinuria, acute recurrent, autosomal recessive |
| LPIN2 | Majeed syndrome |
| LPL | Lipoprotein lipase deficiency |
| LRAT | Leber congenital amaurosis type 14 |
| LRBA | Immunodeficiency, common variable, 8 with autoimmunity |
| LRIG2 | Urofacial syndrome 2 |
| LRIT3 | Night blindness, congenital stationary (complete), 1F, autosomal recessive |
| LRMDA | Albinism, oculocutaneous, type 7 |
| LRP2 | Donnai-Barrow syndrome |
| LRP4 | Cenani-Lenz syndactyly syndrome |
| LRP5 | Osteoporosis-pseudoglioma syndrome |
| LRPAP1 | Myopia, type 23 autosomal recessive |
| LRPPRC | Mitochondrial complex IV deficiency, nuclear type 5, (French-Canadian) |
| LRRC6 | Ciliary dyskinesia, primary, 19 |
| LRSAM1 | Charcot-Marie-Tooth disease, axonal, type 2P |
| LRTOMT | Deafness, autosomal recessive, type 63 |
| LSS | Cataract 44 |
| LTBP2 | Microspherophakia and/or megalocornea, with ectopia lentis and with or without secondary glaucoma |
| LTBP3 | Dental anomalies and short stature |
| LTBP4 | Cutis laxa, autosomal recessive, type 1C |
| LYRM7 | Mitochondrial complex III deficiency, nuclear type 8 |
| LYST | Chediak-Higashi syndrome |
| LZTFL1 | Bardet-Biedl syndrome, type 17 |
| LZTR1 | Noonan syndrome, type 2 |
| MAG | Spastic paraplegia, type 75 autosomal recessive |
| MAGI2 | Nephrotic syndrome, type 15 |
| MAGT1 | Congenital disorder of glycosylation, type Icc |
| MAK | Retinitis pigmentosa type 62 |
| MALT1 | Immunodeficiency, type 12 |
| MAN1B1 | intellectual developmental disorder, autosomal recessive, type 15 |
| MAN2B1 | Alpha-mannosidosis |
| MANBA | Mannosidosis, beta |
| MAP3K20 | Centronuclear myopathy, type 6 with fiber-type disproportion |
| MAPKBP1 | Nephronophthisis 20 |
| MAPT | Supranuclear palsy, progressive atypical (parkinsonism syndrome) |
| MARS1 | Interstitial lung and liver disease |
| MARS2 | Spastic ataxia, type 3 autosomal recessive |
| MARVELD2 | Deafness, autosomal recessive, type 49 |
| MASP1 | 3MC syndrome 1 |
| MAT1A | Methionine adenosyltransferase deficiency, autosomal recessive |
| MATN3 | Spondyloepimetaphyseal dysplasia, Borochowitz-Cormier-Daire type |
| MBOAT7 | intellectual developmental disorder, autosomal recessive 57 |
| MBTPS2 | Olmsted syndrome, X-linked |
| MC2R | Glucocorticoid deficiency, due to ACTH unresponsiveness |
| MCCC1 | 3-Methylcrotonyl-CoA carboxylase deficiency, type 1 |
| MCCC2 | 3-Methylcrotonyl-CoA carboxylase deficiency, type 2 |
| MCEE | Methylmalonyl-CoA epimerase deficiency |
| MCFD2 | Combined deficiency of factor V and factor VIII, type 2 |
| MCIDAS | Ciliary dyskinesia, primary, type 42 |
| MCM3AP | Peripheral neuropathy, autosomal recessive, with or without impaired intellectual development |
| MCM4 | Immunodeficiency, type 54 |
| MCM9 | Ovarian dysgenesis 4 |
| MCOLN1 | Mucolipidosis type 4 |
| MCPH1 | Microcephaly type 1 primary, autosomal recessive |
| MDH2 | Epileptic encephalopathy, early infantile, 51 |
| MECP2 | Encephalopathy, neonatal severe; Rett syndrome |
| MECR | Dystonia, childhood-onset, with optic atrophy and basal ganglia abnormalities |
| MED17 | Microcephaly, postnatal progressive, with seizures and brain atrophy |
| MED23 | intellectual developmental disorder, autosomal recessive, type 18 |
| MED25 | Basel-Vanagait-Smirin-Yosef syndrome |
| MEFV | Familial Mediterranean fever |
| MEGF10 | Myopathy, areflexia, respiratory distress, and dysphagia, early-onset |
| MEGF8 | Carpenter syndrome, type 2 |
| MEOX1 | Klippel-Feil syndrome 2 |
| MERTK | Retinitis pigmentosa type 38 |
| MESP2 | Spondylocostal dysostosis, type 2 autosomal recessive |
| METTL23 | intellectual developmental disorder, autosomal recessive 44 |
| MFF | Encephalopathy due to defective mitochondrial and peroxisomal fission, type 2 |
| MFN2 | Charcot-Marie-Tooth disease, axonal, type 2A2B |
| MFRP | Microphthalmia, isolated type 5 |
| MFSD2A | Microcephaly 15 primary, autosomal recessive |
| MFSD8 | Ceroid lipofuscinosis, neuronal, type 7 |
| MGAT2 | Congenital disorder of glycosylation, type 2a |
| MGME1 | Mitochondrial DNA depletion syndrome 11 |
| MGP | Keutel syndrome |
| MICU1 | Myopathy with extrapyramidal signs |
| MID1 | Opitz GBBB syndrome, type 1 |
| MIPEP | Combined oxidative phosphorylation deficiency 31 |
| MITF | COMMAD syndrome |
| MKKS | Bardet-Biedl syndrome type 6 |
| MKS1 | Bardet-Biedl syndrome type 13; Meckel syndrome, type 1; Joubert syndrome, type 28 |
| MLC1 | Megalencephalic leukoencephalopathy with subcortical cysts |
| MLPH | Griscelli syndrome, type 3 |
| MLYCD | Malonyl-CoA decarboxylase deficiency |
| MMAA | Methylmalonic aciduria, vitamin B12-responsive |
| MMAB | Methylmalonic aciduria, vitamin B12-responsive, type cblB |
| MMACHC | Methylmalonic aciduria and homocystinuria, cblC type |
| MMADHC | Homocystinuria, cblD type, variant 1 |
| MME | Charcot-Marie-Tooth disease, axonal, type 2T |
| MMP13 | Metaphyseal dysplasia, Spahr type |
| MMP2 | Multicentric osteolysis, nodulosis, and arthropathy (MONA) |
| MMP20 | Amelogenesis imperfecta, type 2A2 (hypomaturation type) |
| MMP21 | Heterotaxy, visceral, 7 autosomal |
| MMUT | Methylmalonic aciduria, mut(0) type |
| MOCOS | Xanthinuria, type 2 |
| MOCS1 | Molybdenum cofactor deficiency A |
| MOCS2 | Molybdenum cofactor deficiency B |
| MOGS | Congenital disorder of glycosylation, type 2B |
| MPC1 | Mitochondrial pyruvate carrier deficiency |
| MPDU1 | Congenital disorder of glycosylation, type 1F |
| MPDZ | Hydrocephalus, congenital, type 2 with or without brain or eye anomalies |
| MPI | Congenital disorder of glycosylation, type 1B |
| MPIG6B | Thrombocytopenia, anemia, and myelofibrosis |
| MPL | Thrombocytopenia, congenital amegakaryocytic |
| MPLKIP | Trichothiodystrophy, type 4 nonphotosensitive |
| MPO | Myeloperoxidase deficiency |
| MPV17 | Mitochondrial DNA depletion syndrome type 6 (hepatocerebral); Charcot-Marie-Tooth disease, axonal, type 2EE |
| MPZ | Dejerine-Sottas disease |
| MRAP | Glucocorticoid deficiency, type 2 |
| MRE11 | Ataxia-telangiectasia-like disorder 1 |
| MRPS16 | Combined oxidative phosphorylation deficiency 2 |
| MRPS22 | Combined oxidative phosphorylation deficiency type 5 |
| MRPS34 | Combined oxidative phosphorylation deficiency 32 |
| MSH3 | Familial adenomatous polyposis, type 4 |
| MSMO1 | Microcephaly, congenital cataract, and psoriasiform dermatitis |
| MSRB3 | Deafness, autosomal recessive, type 74 |
| MSTO1 | Myopathy, mitochondrial, and ataxia |
| MTFMT | Combined oxidative phosphorylation deficiency 15 |
| MTHFD1 | Combined immunodeficiency and megaloblastic anemia with or without hyperhomocysteinemia |
| MTHFR | Homocystinuria due to MTHFR deficiency |
| MTM1 | Myotubular myopathy, X-linked |
| MTMR2 | Charcot-Marie-Tooth disease, type 4B1 |
| MTO1 | Combined oxidative phosphorylation deficiency 10 |
| MTR | Homocystinuria-megaloblastic anemia, cblG complementation type |
| MTRR | Homocystinuria-megaloblastic anemia, cbl E type |
| MTTP | Abetalipoproteinemia |
| MUSK | Fetal akinesia deformation sequence, type 1; Myasthenic syndrome, congenital, type 9 associated with acetylcholine receptor deficiency |
| MUTYH | Adenomas, multiple colorectal |
| MVK | Mevalonic aciduria |
| MYBPC1 | Lethal congenital contracture syndrome, type 4 |
| MYD88 | Immunodeficiency, type 68 |
| MYH2 | Proximal myopathy and ophthalmoplegia |
| MYMK | Carey-Fineman-Ziter syndrome |
| MYO15A | Deafness, autosomal recessive, type 3 |
| MYO18B | Klippel-Feil syndrome, type 4 autosomal recessive, with myopathy and facial dysmorphism |
| MYO1E | Glomerulosclerosis, focal segmental, 6 |
| MYO3A | Deafness, autosomal recessive, type 30 |
| MYO5A | Griscelli syndrome, type 1 |
| MYO5B | Microvillus inclusion disease |
| MYO6 | Deafness, autosomal recessive, type 37 |
| MYO7A | Usher syndrome, type 1B; Deafness, autosomal recessive, type 2 |
| MYPN | Nemaline myopathy, type 11 autosomal recessive |
| NADK2 | 2,4-dienoyl-CoA reductase deficiency |
| NAGA | Schindler disease, type I |
| NAGLU | Mucopolysaccharidosis, type 3B (Sanfilippo B) |
| NAGS | N-acetylglutamate synthase deficiency |
| NALCN | Hypotonia, infantile, with psychomotor retardation and characteristic facies 1 |
| NANS | Spondyloepimetaphyseal dysplasia, Camera-Genevieve type |
| NARS2 | Combined oxidative phosphorylation deficiency 24 |
| NAXE | Encephalopathy, progressive, early-onset, with brain edema and/or leukoencephalopathy |
| NBAS | Infantile liver failure syndrome, type 2; Short stature, optic nerve atrophy, and Pelger-Huet anomaly |
| NBEAL2 | Gray platelet syndrome |
| NBN | Nijmegen breakage syndrome |
| NCAPD3 | Microcephaly 22 primary, autosomal recessive |
| NCF1 | Chronic granulomatous disease, type 1 |
| NCF2 | Chronic granulomatous disease, type 2 |
| NCF4 | Chronic granulomatous disease 3 autosomal recessive |
| NDE1 | Lissencephaly, type 4 (with microcephaly) |
| NDP | Norrie disease |
| NDRG1 | Charcot-Marie-Tooth disease, type 4D |
| NDST1 | intellectual developmental disorder, autosomal recessive, type 46 |
| NDUFA1 | Mitochondrial complex I deficiency, nuclear type 12 |
| NDUFA10 | Mitochondrial complex I deficiency, nuclear type 22 |
| NDUFA11 | Mitochondrial complex I deficiency, nuclear type 14 |
| NDUFA12 | Mitochondrial complex I deficiency, nuclear type 23 |
| NDUFA2 | Mitochondrial complex I deficiency, nuclear type 13 |
| NDUFA9 | Mitochondrial complex I deficiency, nuclear type 26 |
| NDUFAF1 | Mitochondrial complex I deficiency, nuclear type 11 |
| NDUFAF2 | Mitochondrial complex I deficiency, nuclear type 10 |
| NDUFAF3 | Mitochondrial complex I deficiency, nuclear type 18 |
| NDUFAF5 | Mitochondrial complex I deficiency, nuclear type 16 |
| NDUFAF6 | Mitochondrial complex I deficiency, nuclear type 17 |
| NDUFB3 | Mitochondrial complex I deficiency, nuclear type 25 |
| NDUFB9 | Mitochondrial complex I deficiency, nuclear type 24 |
| NDUFS1 | Mitochondrial complex I deficiency, nuclear type 5 |
| NDUFS2 | Mitochondrial complex I deficiency, nuclear type 6 |
| NDUFS3 | Mitochondrial complex I deficiency, nuclear type 8 |
| NDUFS4 | Mitochondrial complex I deficiency, nuclear type 1 |
| NDUFS6 | Mitochondrial complex I deficiency, nuclear type 9 |
| NDUFS7 | Mitochondrial complex I deficiency, nuclear type 3 |
| NDUFS8 | Mitochondrial complex I deficiency, nuclear type 2 |
| NDUFV1 | Mitochondrial complex I deficiency, nuclear type 4 |
| NDUFV2 | Mitochondrial complex I deficiency, nuclear type 7 |
| NEB | Nemaline myopathy type 2 |
| NECTIN1 | Cleft lip/palate-ectodermal dysplasia syndrome; Orofacial cleft 7 |
| NECTIN4 | Ectodermal dysplasia-syndactyly syndrome, type 1 |
| NEFL | Charcot-Marie-Tooth disease, type 1F |
| NEK1 | Short-rib thoracic dysplasia, type 6 with or without polydactyly |
| NEK8 | Renal-hepatic-pancreatic dysplasia, type 2 |
| NEK9 | Lethal congenital contracture syndrome 10 |
| NEU1 | Sialidosis, type 1 and type 2 |
| NEUROG3 | Diarrhea 4 malabsorptive, congenital |
| NFU1 | Multiple mitochondrial dysfunctions syndrome 1 |
| NGF | Neuropathy, hereditary sensory and autonomic, type 5 |
| NGLY1 | Congenital disorder of deglycosylation |
| NHEJ1 | Severe combined immunodeficiency with microcephaly, growth retardation, and sensitivity to ionizing radiation |
| NHLRC1 | Epilepsy, progressive myoclonic, type 2B (Lafora) |
| NHP2 | Dyskeratosis congenita, autosomal recessive type 2 |
| NHS | Nance-Horan syndrome; Cataract 40, X-Linked |
| NIN | Seckel syndrome, type 7 |
| NIPAL4 | Ichthyosis, congenital, autosomal recessive, type 6 |
| NKX2-6 | Conotruncal heart malformations |
| NKX3-2 | Spondylo-megaepiphyseal-metaphyseal dysplasia |
| NKX6-2 | Spastic ataxia 8 autosomal recessive, with hypomyelinating leukodystrophy |
| NLRP1 | Autoinflammation with arthritis and dyskeratosis |
| NLRP7 | Hydatidiform mole, recurrent, type 1 |
| NME8 | Ciliary dyskinesia, primary, type 6 |
| NMNAT1 | Leber congenital amaurosis type 9 |
| NNT | Glucocorticoid deficiency 4 with or without mineralocorticoid deficiency |
| NONO | Intellectual developmental disorder, X-linked syndromic 34 |
| NOP10 | Dyskeratosis congenita, autosomal recessive type 1 |
| NPC1 | Niemann-Pick disease, type C1 |
| NPC2 | Niemann-pick disease, type C2 |
| NPHP1 | Joubert syndrome type 4 |
| NPHP3 | Meckel syndrome type 7 |
| NPHP4 | Nephronophthisis type 4 |
| NPHS1 | Nephrotic syndrome, type 1 |
| NPHS2 | Nephrotic syndrome, type 2 |
| NPR2 | Acromesomelic dysplasia, Maroteaux type |
| NR0B1 | Adrenal hypoplasia, congenital |
| NR1H4 | Cholestasis, progressive familial intrahepatic, type 5 |
| NR2E3 | Enhanced S-cone syndrome (Goldmann-Favre); Retinitis pigmentosa, type 37 |
| NRL | Retinal degeneration, autosomal recessive, clumped pigment type |
| NRXN1 | Pitt-Hopkins-like syndrome, type 2 |
| NSDHL | CHILD syndrome; CK syndrome |
| NSMCE2 | Seckel syndrome, type 10 |
| NSUN2 | Intellectual developmental disorder, autosomal recessive 5 |
| NT5C2 | Spastic paraplegia, type 45 autosomal recessive |
| NT5C3A | Anemia, hemolytic, due to UMPH1 deficiency |
| NT5E | Calcification of joints and arteries |
| NTHL1 | Familial adenomatous polyposis, type 3 |
| NTRK1 | Insensitivity to pain, congenital, with anhidrosis |
| NUBPL | Mitochondrial complex I deficiency, nuclear type 21 |
| NUP107 | Nephrotic syndrome, type 11 |
| NUP62 | Striatonigral degeneration, infantile |
| NUP93 | Nephrotic syndrome, type 12 |
| NYX | Night blindness, congenital stationary (complete), 1A, X-linked |
| OAT | Gyrate atrophy of choroid and retina |
| OBSL1 | 3M syndrome 2 |
| OCA2 | Oculocutaneous albinism type 2 |
| OCLN | Pseudo-TORCH syndrome, type 1 |
| OCRL | Lowe Syndrome; Dent disease type 2 |
| ODAD1 | Ciliary dyskinesia, primary, type 20 |
| ODAD2 | Ciliary dyskinesia, primary, type 23 |
| ODAD3 | Ciliary dyskinesia, primary, type 30 |
| OFD1 | Joubert syndrome 10; Simpson-Golabi-Behmel syndrome, type 2 |
| OPA1 | Behr syndrome |
| OPA3 | 3-methylglutaconic aciduria, type 3 |
| OPHN1 | Intellectual developmental disorder, X-linked syndromic, Billuart type |
| ORAI1 | Immunodeficiency, type 9 |
| ORC1 | Meier-Gorlin syndrome, type 1 |
| ORC4 | Meier-Gorlin syndrome, type 2 |
| ORC6 | Meier-Gorlin syndrome, type 3 |
| OSGEP | Galloway-Mowat syndrome 3 |
| OSTM1 | Osteopetrosis, autosomal recessive type 5 |
| OTC | Ornithine transcarbamylase deficiency |
| OTOA | Deafness, autosomal recessive, type 22 |
| OTOF | Deafness, autosomal recessive, type 9 |
| OTOG | Deafness, autosomal recessive, type 18B |
| OTOGL | Deafness, autosomal recessive, type 84B |
| OTUD6B | Intellectual developmental disorder with dysmorphic facies, seizures, and distal limb anomalies |
| OTULIN | Autoinflammation, panniculitis, and dermatosis syndrome |
| OXCT1 | Succinyl CoA:3-oxoacid CoA transferase deficiency |
| P2RY12 | Bleeding disorder, platelet-type, type 8 |
| P3H1 | Osteogenesis imperfecta, type 8 |
| P3H2 | Myopia, high, with cataract and vitreoretinal degeneration |
| PADI6 | Preimplantation embryonic lethality 2 |
| PAH | Phenylketonuria |
| PAK3 | Intellectual developmental disorder, X-linked 30 |
| PALB2 | Fanconi anemia, complementation group N |
| PAM16 | Spondylometaphyseal dysplasia, Megarbane-Dagher-Melike type |
| PANK2 | Neurodegeneration with brain iron accumulation type 1 |
| PAPSS2 | Brachyolmia, type 4 with mild epiphyseal and metaphyseal changes |
| PARK7 | Parkinson disease, type 7 autosomal recessive, early-onset |
| PARN | Dyskeratosis congenita, autosomal recessive 6 |
| PATL2 | Oocyte maturation defect 4 |
| PAX7 | Rhabdomyosarcoma 2 alveolar |
| PC | Pyruvate carboxylase deficiency |
| PCARE | Retinitis pigmentosa, type 54 |
| PCBD1 | Hyperphenylalaninemia, BH4-deficient, type D |
| PCCA | Propionic acidemia |
| PCCB | Propionic acidemia |
| PCDH12 | Microcephaly, seizures, spasticity, and brain calcification |
| PCDH15 | Deafness, autosomal recessive, type 23; Usher syndrome, type 1D/F digenic |
| PCDH19 | Developmental and epileptic encephalopathy 9 |
| PCK2 | PEPCK deficiency, mitochondrial |
| PCNT | Microcephalic osteodysplastic primordial dwarfism, type 2 |
| PCSK1 | Obesity with impaired prohormone processing |
| PCYT1A | Spondylometaphyseal dysplasia with cone-rod dystrophy |
| PDE10A | Dyskinesia, limb and orofacial, infantile-onset |
| PDE6A | Retinitis pigmentosa type 43 |
| PDE6B | Retinitis pigmentosa type 40 |
| PDE6C | Cone dystrophy type 4 |
| PDE6G | Retinitis pigmentosa type 57 |
| PDE6H | Retinal cone dystrophy 3 and achromatopsia 6 |
| PDHA1 | Pyruvate dehydrogenase E1-alpha deficiency |
| PDHB | Pyruvate dehydrogenase E1-beta deficiency |
| PDHX | Lacticacidemia due to PDX1 deficiency |
| PDP1 | Pyruvate dehydrogenase phosphatase deficiency |
| PDSS1 | Coenzyme Q10 deficiency, primary, type 2 |
| PDSS2 | Coenzyme Q10 deficiency, primary, type 3 |
| PDX1 | Pancreatic agenesis type 1 |
| PDXK | Neuropathy, hereditary motor and sensory, type VIC, with optic atrophy |
| PEPD | Prolidase deficiency |
| PET100 | Mitochondrial complex IV deficiency, nuclear type 12 |
| PEX1 | Heimler syndrome type 1 |
| PEX10 | Peroxisome biogenesis disorder, type 6A (Zellweger syndrome); Peroxisome biogenesis disorder, type 6B |
| PEX11B | Peroxisome biogenesis disorder 14B |
| PEX12 | Peroxisome biogenesis disorder type 3A (Zellweger) |
| PEX13 | Peroxisome biogenesis disorder, type 11A (Zellweger syndrome); Peroxisome biogenesis disorder, type 11B |
| PEX14 | Peroxisome biogenesis disorder, type 13A (Zellweger syndrome) |
| PEX16 | Peroxisome biogenesis disorder, type 8A (Zellweger syndrome); Peroxisome biogenesis disorder, type 8B |
| PEX19 | Peroxisome biogenesis disorder, type 12A (Zellweger syndrome) |
| PEX2 | Peroxisome biogenesis disorder type 5A (Zellweger) |
| PEX26 | Peroxisome biogenesis disorder type 7A (Zellweger) |
| PEX3 | Peroxisome biogenesis disorder, type 10A (Zellweger syndrome) |
| PEX5 | Peroxisome biogenesis disorder type 2A (Zellweger) |
| PEX6 | Peroxisome biogenesis disorder, type 4A (Zellweger syndrome); Peroxisome biogenesis disorder, type 4B; Heimler syndrome 2 |
| PEX7 | Rhizomelic chondrodysplasia punctata, type 1 |
| PFKM | Glycogen storage disease, type 7 |
| PGAM2 | Glycogen storage disease X |
| PGAP1 | intellectual developmental disorder, autosomal recessive 42 |
| PGAP2 | Hyperphosphatasia with impaired intellectual development syndrome 3 |
| PGAP3 | Hyperphosphatasia with impaired intellectual development syndrome 4 |
| PGK1 | Phosphoglycerate kinase 1 deficiency |
| PGM1 | Congenital disorder of glycosylation, type 1t |
| PGM3 | Immunodeficiency, type 23 |
| PHF8 | Intellectual developmental disorder, X-linked syndromic, Siderius type |
| PHGDH | Neu-Laxova syndrome, type 1; Phosphoglycerate dehydrogenase deficiency |
| PHKA1 | Muscle glycogenosis |
| PHKA2 | Glycogen storage disease, type IXa1/2 |
| PHKB | Glycogen storage disease, type 9B |
| PHKG2 | Glycogen storage disease type 9c |
| PHOX2A | Fibrosis of extraocular muscles, congenital, 2 |
| PHYH | Refsum disease |
| PI4KA | Polymicrogyria, perisylvian, with cerebellar hypoplasia and arthrogryposis |
| PIBF1 | Joubert syndrome 33 |
| PIEZO1 | Lymphedema, hereditary, type 3 |
| PIEZO2 | Arthrogryposis, distal, with impaired proprioception and touch |
| PIGC | Glycosylphosphatidylinositol biosynthesis defect 16 |
| PIGG | intellectual developmental disorder, autosomal recessive 53 |
| PIGL | Zunich neuroectodermal syndrome |
| PIGM | Glycosylphosphatidylinositol deficiency |
| PIGN | Multiple congenital anomalies-hypotonia-seizures syndrome, type 1 |
| PIGO | Hyperphosphatasia with impaired intellectual development syndrome 2 |
| PIGT | Multiple congenital anomalies-hypotonia-seizures syndrome 3 |
| PIGV | Hyperphosphatasia with impaired intellectual development syndrome 1 |
| PIGW | Glycosylphosphatidylinositol biosynthesis defect 11 |
| PIGY | Hyperphosphatasia with impaired intellectual development syndrome 6 |
| PINK1 | Parkinson disease, type 6 early onset |
| PIP5K1C | Lethal congenital contractural syndrome, type 3 |
| PJVK | Deafness, autosomal recessive, type 59 |
| PKD1L1 | Heterotaxy, visceral, 8 autosomal |
| PKHD1 | Polycystic kidney disease type 4 |
| PKLR | Pyruvate kinase deficiency |
| PKP1 | Ectodermal dysplasia/skin fragility syndrome |
| PLA2G6 | Infantile neuroaxonal dystrophy type 1 |
| PLAA | Neurodevelopmental disorder with progressive microcephaly, spasticity, and brain anomalies |
| PLCB1 | Epileptic encephalopathy, early infantile, type 12 |
| PLCB4 | Auriculocondylar syndrome, type 2 |
| PLCD1 | Nail disorder, nonsyndromic congenital, type 3 (leukonychia) |
| PLCE1 | Nephrotic syndrome, type 3 |
| PLD1 | Cardiac valvular defect, developmental |
| PLEC | Epidermolysis bullosa simplex with muscular dystrophy |
| PLEKHG5 | Charcot-Marie-Tooth disease, recessive intermediate, type C |
| PLG | Plasminogen deficiency, type I |
| PLK4 | Microcephaly and chorioretinopathy, autosomal recessive, 2 |
| PLOD1 | Ehlers-Danlos syndrome, kyphoscoliotic type, 1 |
| PLOD2 | Bruck syndrome 2 |
| PLOD3 | Lysyl hydroxylase 3 deficiency |
| PLP1 | Pelizaeus-Merzbacher disease |
| PLPBP | Epilepsy, early-onset, vitamin B6-dependent |
| PMM2 | Congenital disorder of glycosylation, type 1A |
| PMP22 | Dejerine-Sottas disease |
| PMPCA | Spinocerebellar ataxia, autosomal recessive, type 2 |
| PMPCB | Multiple mitochondrial dysfunctions syndrome 6 |
| PNKP | Ataxia-oculomotor apraxia, type 4; Microcephaly, seizures, and developmental delay |
| PNP | Immunodeficiency due to purine nucleoside phosphorylase deficiency |
| PNPLA1 | Ichthyosis, congenital, autosomal recessive, type 10 |
| PNPLA2 | Neutral lipid storage disease with myopathy |
| PNPLA6 | Boucher-Neuhauser syndrome; Oliver-McFarlane syndrome; Spastic paraplegia, type 39 autosomal recessive |
| PNPO | Pyridoxamine 5'-phosphate oxidase deficiency |
| PNPT1 | Combined oxidative phosphorylation deficiency 13 |
| POC1A | Short stature, onychodysplasia, facial dysmorphism, and hypotrichosis |
| POC1B | Cone-rod dystrophy 20 |
| POLE | FILS syndrome |
| POLG | POLG-related disorders |
| POLH | Xeroderma pigmentosum, variant type |
| POLR1C | Leukodystrophy, hypomyelinating, type 11; Treacher Collins syndrome 3 |
| POLR1D | Treacher Collins syndrome, type 2 |
| POLR3A | Leukodystrophy, hypomyelinating, type 7 |
| POLR3B | Leukodystrophy, hypomyelinating, type 8 |
| POMC | Obesity, adrenal insufficiency, and red hair due to POMC deficiency |
| POMGNT1 | Muscular dystrophy-dystroglycanopathy, type 3A (Walker-Warburg syndrome); Type 3B; Type 3C (limb-girdle muscular dystrophy, type 15 [LGMDR15]) |
| POMGNT2 | Muscular dystrophy-dystroglycanopathy, type 8A (Walker-Warburg syndrome); Type 8C (limb-girdle muscular dystrophy, type 24 [LGMD R24]) |
| POMK | Muscular dystrophy-dystroglycanopathy (congenital with brain and eye anomalies), type A, 12 |
| POMP | Keratosis linearis with ichthyosis congenita and sclerosing keratoderma |
| POMT1 | Muscular dystrophy-dystroglycanopathy, type 1A (Walker-Warburg syndrome); Type 1B; Type 1C (limb-girdle muscular dystrophy, type 11 [LGMD R11]) muscular dystrophy, type 11 [LGMD R11]) |
| POMT2 | Muscular dystrophy-dystroglycanopathy, type 2A (Walker-Warburg syndrome); Type 2B; Type 2C (limb-girdle muscular dystrophy, type 14 [LGMD R14]) muscular dystrophy, type 14 [LGMD R14]) |
| POP1 | Anauxetic dysplasia, type 2 |
| POR | Antley-Bixler syndrome with genital anomalies and disordered steroidogenesis |
| POU1F1 | Pituitary hormone deficiency, combined, type 1 |
| POU3F4 | Deafness, X-linked, type 2 |
| PPA2 | Sudden cardiac failure, infantile |
| PPIB | Osteogenesis imperfecta, type 9 |
| PPM1K | Maple syrup urine disease, mild variant |
| PPP1R15B | Microcephaly, short stature, and impaired glucose metabolism 2 |
| PPT1 | Ceroid lipofuscinosis, neuronal, type 1 |
| PQBP1 | Renpenning syndrome |
| PRCD | Retinitis pigmentosa, type 36 |
| PRDM12 | Neuropathy, hereditary sensory and autonomic, type VIII |
| PRDM5 | Brittle cornea syndrome, type 2 |
| PRDX1 | Methylmalonic aciduria and homocystinuria, cblC type, digenic |
| PREPL | Myasthenic syndrome, congenital, type 22 |
| PRF1 | Hemophagocytic lymphohistiocytosis, familial, type 2 |
| PRG4 | Camptodactyly-arthropathy-coxa vara-pericarditis syndrome |
| PRICKLE1 | Epilepsy, progressive myoclonic, type 1B |
| PRKCD | Autoimmune lymphoproliferative syndrome, type 3 |
| PRKDC | Immunodeficiency 26, with or without neurologic abnormalities |
| PRKN | Parkinson disease, type 2 juvenile |
| PRKRA | Dystonia, type 16 |
| PRMT7 | Short stature, brachydactyly, intellectual developmental disability, and seizures |
| PROC | Thrombophilia due to protein C deficiency, autosomal recessive |
| PRODH | Hyperprolinemia, type 1 |
| PROM1 | Retinitis pigmentosa, type 41 |
| PROP1 | Pituitary hormone deficiency, combined, type 2 |
| PROS1 | Thrombophilia due to protein S deficiency, autosomal recessive |
| PRPH2 | Leber congenital amaurosis 18; Retinitis punctata albescens |
| PRPS1 | Phosphoribosylpyrophosphate synthetase (PRS) deficiency |
| PRRX1 | Agnathia-otocephaly complex |
| PRSS1 | Trypsinogen deficiency |
| PRSS12 | Intellectual developmental disorder, autosomal recessive 1 |
| PRSS56 | Microphthalmia, isolated, type 6 |
| PRUNE1 | Neurodevelopmental disorder with microcephaly, hypotonia, and variable brain anomalies |
| PRX | Charcot-Marie-Tooth disease, type 4F |
| PSAP | Combined SAP deficiency |
| PSAT1 | Neu-Laxova syndrome, type 2 |
| PSMB8 | Autoinflammation, lipodystrophy, and dermatosis syndrome |
| PSMC3IP | Ovarian dysgenesis 3 |
| PSPH | Phosphoserine phosphatase deficiency |
| PTF1A | Pancreatic agenesis 2 |
| PTH | Hypoparathyroidism, familial isolated, type 1 |
| PTH1R | Chondrodysplasia, Blomstrand type; Eiken syndrome |
| PTPN23 | Neurodevelopmental disorder and structural brain anomalies with or without seizures and spasticity |
| PTPRC | Severe combined immunodeficiency, T cell-negative, B-cell/natural killer-cell positive |
| PTPRO | Nephrotic syndrome, type 6 |
| PTPRQ | Deafness, autosomal recessive, type 84A |
| PTRH2 | Infantile-onset multisystem neurologic, endocrine, and pancreatic disease |
| PTS | Hyperphenylalaninemia, BH4-deficient, type A |
| PUS1 | Myopathy, lactic acidosis, and sideroblastic anemia, type 1 |
| PXDN | Anterior segment dysgenesis, type 7 with sclerocornea |
| PYCR1 | Cutis laxa, autosomal recessive, type 2B |
| PYCR2 | Leukodystrophy, hypomyelinating, type 10 |
| PYGL | Glycogen storage disease, type 6 |
| PYGM | McArdle disease |
| PYROXD1 | Myopathy, myofibrillar, type 8 |
| QARS1 | Microcephaly, progressive, seizures, and cerebral and cerebellar atrophy |
| QDPR | Hyperphenylalaninemia, BH4-deficient, type C |
| RAB18 | Warburg micro syndrome, type 3 |
| RAB23 | Carpenter syndrome |
| RAB27A | Griscelli syndrome, type 2 |
| RAB28 | Cone-rod dystrophy 18 |
| RAB33B | Smith-McCort dysplasia 2 |
| RAB39B | Intellectual developmental disorder, X-linked 72 |
| RAB3GAP1 | Warburg micro syndrome; Martsolf syndrome |
| RAB3GAP2 | Warburg micro syndrome; Martsolf syndrome |
| RAD50 | Nijmegen breakage syndrome-like disorder |
| RAD51C | Fanconi anemia, complementation group O |
| RAG1 | Omenn syndrome; Severe combined immunodeficiency, B cell-negative |
| RAG2 | Omenn syndrome; Severe combined immunodeficiency, B cell-negative |
| RAPSN | Fetal akinesia deformation sequence, type 2; Myasthenic syndrome, congenital, type 11 associated with AChR deficiency |
| RARB | Microphthalmia, syndromic 12 |
| RARS1 | Leukodystrophy, hypomyelinating, type 9 |
| RARS2 | Pontocerebellar hypoplasia, type 6 |
| RASGRP1 | Immunodeficiency, type 64 |
| RAX | Microphthalmia, syndromic 16 |
| RBBP8 | Jawad syndrome; Seckel syndrome, type 2 |
| RBCK1 | Polyglucosan body myopathy 1 with or without immunodeficiency |
| RBM8A | Thrombocytopenia-absent radius syndrome |
| RBP3 | Retinitis pigmentosa 66 |
| RBP4 | Retinal dystrophy, iris coloboma, and comedogenic acne syndrome |
| RCBTB1 | Retinal dystrophy with or without extraocular anomalies |
| RD3 | Leber congenital amaurosis, type 12 |
| RDH12 | Leber congenital amaurosis, type 13 |
| RDH5 | Fundus albipunctatus |
| RDX | Deafness, autosomal recessive, type 24 |
| RECQL4 | Baller-Gerold syndrome; RAPADILINO syndrome; Rothmund-Thomson syndrome |
| REEP6 | Retinitis pigmentosa 77 |
| RELN | Lissencephaly 2 (Norman-Roberts type) |
| REN | Renal tubular dysgenesis |
| RETREG1 | Neuropathy, hereditary sensory and autonomic, type 2B |
| RFT1 | Congenital disorder of glycosylation, type In |
| RFX5 | Bare lymphocyte syndrome, type 2 |
| RFX6 | Mitchell-Riley syndrome |
| RFXANK | Bare lymphocyte syndrome, type 2 complementation group B |
| RFXAP | Bare lymphocyte syndrome, type 2 |
| RHAG | Anemia, hemolytic, Rh-null, regulator type |
| RHO | Retinitis pigmentosa, type 4; Retinitis punctata albescens |
| RIN2 | Macs syndrome |
| RIPK4 | Popliteal pterygium syndrome, Bartsocas-Papas type |
| RIPOR2 | Deafness, autosomal recessive, type 104 |
| RLBP1 | Bothnia retinal dystrophy; Fundus albipunctatus |
| RMND1 | Combined oxidative phosphorylation deficiency 11 |
| RMRP | Anauxetic dysplasia 1 |
| RNASEH1 | Progressive external ophthalmoplegia with mitochondrial DNA deletions, autosomal recessive 2 |
| RNASEH2A | Aicardi-Goutieres syndrome, type 4 |
| RNASEH2B | Aicardi-Goutieres syndrome, type 2 |
| RNASEH2C | Aicardi-Goutieres syndrome, type 3 |
| RNASET2 | Leukoencephalopathy, cystic, without megalencephaly |
| RNF168 | RIDDLE syndrome |
| RNF216 | Gordon Holmes syndrome |
| ROBO3 | Gaze palsy, familial horizontal, with progressive scoliosis, type 1 |
| ROGDI | Kohlschutter-Tonz syndrome |
| ROM1 | Retinitis pigmentosa, type 7 digenic |
| ROR2 | Robinow syndrome, autosomal recessive |
| RORC | Immunodeficiency, type 42 |
| RP1 | Retinitis pigmentosa, type 1 |
| RP2 | Retinitis pigmentosa, type 2 X-linked |
| RPE65 | RPE65-related Leber congenital amaurosis/early-onset severe retinal dystrophy |
| RPGR | Retinitis pigmentosa, type 3 X-linked; Cone-rod dystrophy, X-linked, 1 |
| RPGRIP1 | Leber congenital amaurosis, type 6 |
| RPGRIP1L | Joubert syndrome, type 7; Meckel syndrome, type 5; COACH syndrome |
| RPL10 | Intellectual developmental disorder, X-linked syndromic 35 |
| RPS6KA3 | Coffin-Lowry syndrome |
| RRM2B | Mitochondrial DNA depletion syndrome, type 8A (encephalomyopathic type with renal tubulopathy) and type 8B (MNGIE type) |
| RS1 | Retinoschisis |
| RSPH1 | Ciliary dyskinesia, primary, type 24 |
| RSPH3 | Ciliary dyskinesia, primary, type 32 |
| RSPH4A | Ciliary dyskinesia, primary, type 11 |
| RSPH9 | Ciliary dyskinesia, primary, type 12 |
| RSPO4 | Anonychia congenita |
| RSPRY1 | Spondyloepimetaphyseal dysplasia, Faden-Alkuraya type |
| RTEL1 | Dyskeratosis congenita, autosomal recessive type 5 |
| RTN4IP1 | Optic atrophy 10 with or without ataxia, impaired intellectual development and seizures |
| RTTN | Microcephaly, short stature, and polymicrogyria with seizures |
| RUSC2 | intellectual developmental disorder, autosomal recessive 61 |
| RXYLT1 | Muscular dystrophy-dystroglycanopathy (congenital with brain and eye anomalies), type A, 10 |
| RYR1 | RYR1 related congenitalmyopathy |
| S1PR2 | Deafness, autosomal recessive, type 68 |
| SACS | Spastic ataxia, Charlevoix-Saguenay, type |
| SAG | Oguchi disease, type 1 |
| SAMD9 | Tumoral calcinosis, familial, normophosphatemic |
| SAMHD1 | Aicardi-Goutieres syndrome, type 5 |
| SAR1B | Chylomicron retention disease |
| SARS2 | Hyperuricemia, pulmonary hypertension, renal failure, and alkalosis |
| SBDS | Shwachman-Diamond syndrome |
| SBF1 | Charcot-Marie-Tooth disease, type 4B3 |
| SBF2 | Charcot-Marie-Tooth disease, type 4B2 |
| SC5D | Lathosterolosis |
| SCARB2 | Epilepsy, progressive myoclonic, type 4 with or without renal failure |
| SCARF2 | Van den Ende-Gupta syndrome |
| SCN1B | Epileptic encephalopathy, early infantile, type 52 |
| SCN4A | Myasthenic syndrome, congenital, type 16 |
| SCN9A | Indifference to pain and autosomal recessive hereditary sensory neuropathy type 2D |
| SCNN1A | Pseudohypoaldosteronism, type 1 |
| SCNN1B | Pseudohypoaldosteronism, type 1 |
| SCNN1G | Pseudohypoaldosteronism, type 1 |
| SCO1 | Mitochondrial complex IV deficiency, nuclear type 4 |
| SCO2 | Mitochondrial complex IV deficiency, nuclear type 2 |
| SCYL1 | Spinocerebellar ataxia, autosomal recessive, type 21 |
| SDCCAG8 | Bardet-Biedl syndrome, type 16 |
| SDHA | Mitochondrial respiratory chain complex II deficiency; Leigh syndrome |
| SDHAF1 | Mitochondrial complex II deficiency |
| SDR9C7 | Ichthyosis, congenital, autosomal recessive 13 |
| SEC23A | Craniolenticulosutural dysplasia |
| SEC23B | Dyserythropoietic anemia, congenital, type 2 |
| SEC24D | Cole-Carpenter syndrome 2 |
| SECISBP2 | Thyroid hormone metabolism, abnormal |
| SELENON | Multiminicore disease (rigid spine syndrome) |
| SEMA4A | Cone-rod dystrophy, type 10; Retinitis pigmentosa, type 35 |
| SEPSECS | Pontocerebellar hypoplasia, type 2D |
| SERAC1 | 3-methylglutaconic aciduria with deafness, encephalopathy, and Leigh-like syndrome (MEGDEL) |
| SERPINA1 | Alpha-1 antitrypsin deficiency |
| SERPINB7 | Palmoplantar keratoderma, Nagashima type |
| SERPINB8 | Peeling skin syndrome 5 |
| SERPINC1 | Thrombophilia due to antithrombin III deficiency |
| SERPINE1 | Plasminogen activator inhibitor-1 deficiency |
| SERPINF1 | Osteogenesis imperfecta, type 6 |
| SERPINF2 | Alpha-2-plasmin inhibitor deficiency |
| SERPING1 | Angioedema, hereditary, types 1 and 2 |
| SERPINH1 | Osteogenesis imperfecta, type 10 |
| SETX | Spinocerebellar ataxia, autosomal recessive, type 1 |
| SFRP4 | Pyle disease |
| SFTPB | Surfactant metabolism dysfunction, pulmonary, type 1 |
| SFXN4 | Combined oxidative phosphorylation deficiency 18 |
| SGCA | Limb-girdle muscular dystrophy, type 3 (LGMD R3) |
| SGCB | Limb-girdle muscular dystrophy, type 4 (LGMD R4) |
| SGCD | Limb-girdle muscular dystrophy, type 6 (LGMD R6) |
| SGCG | Limb-girdle muscular dystrophy, type 5 (LGMD R5) |
| SGPL1 | Nephrotic syndrome, type 14 |
| SGSH | Mucopolysaccharidosis, type 3A (Sanfilippo A) |
| SH2D1A | Lymphoproliferative syndrome, X-linked, type 1 |
| SH3PXD2B | Frank-ter Haar syndrome |
| SH3TC2 | Charcot-Marie-Tooth disease, type 4C |
| SI | Sucrase-isomaltase deficiency, congenital |
| SIL1 | Marinesco-Sjogren syndrome |
| SIX6 | Optic disc anomalies with retinal and/or macular dystrophy |
| SKIV2L | Trichohepatoenteric syndrome 2 |
| SLC10A2 | Bile acid malabsorption, primary |
| SLC11A2 | Anemia, hypochromic microcytic, with iron overload 1 |
| SLC12A1 | Bartter syndrome, type 1 |
| SLC12A3 | Gitelman syndrome |
| SLC12A5 | Epileptic encephalopathy, early infantile, 34 |
| SLC12A6 | Agenesis of the corpus callosum with peripheral neuropathy |
| SLC13A5 | Epileptic encephalopathy, early infantile, 25 |
| SLC16A1 | Monocarboxylate transporter 1 deficiency |
| SLC16A2 | Allan-Herndon-Dudley syndrome |
| SLC17A5 | Salla disease |
| SLC18A3 | Myasthenic syndrome, congenital, 21 presynaptic |
| SLC19A2 | Thiamine-responsive megaloblastic anemia syndrome |
| SLC19A3 | Thiamine metabolism dysfunction syndrome, type 2 (biotin- or thiamine-responsive encephalopathy type) |
| SLC1A1 | Dicarboxylic aminoaciduria |
| SLC1A4 | Spastic tetraplegia, thin corpus callosum, and progressive microcephaly |
| SLC22A12 | Hypouricemia, renal |
| SLC22A5 | Carnitine deficiency, systemic primary |
| SLC24A1 | Night blindness, congenital stationary (complete), type 1D, autosomal recessive |
| SLC24A4 | Amelogenesis imperfecta, type IIA5 |
| SLC24A5 | Albinism, oculocutaneous, type 6 |
| SLC25A1 | Combined D-2- and L-2-hydroxyglutaric aciduria |
| SLC25A12 | Epileptic encephalopathy, early infantile, type 39 |
| SLC25A13 | Citrullinemia, type 2 neonatal-onset; Citrullinemia, type 2 adult-onset |
| SLC25A15 | Hyperornithinemia-hyperammonemia-homocitrullinemia syndrome |
| SLC25A19 | Microcephaly, Amish type; Thiamine metabolism dysfunction syndrome 4 (progressive polyneuropathy type) |
| SLC25A20 | Carnitine-acylcarnitine translocase deficiency |
| SLC25A22 | Developmental and epileptic encephalopathy 3 |
| SLC25A26 | Combined oxidative phosphorylation deficiency 28 |
| SLC25A3 | Mitochondrial phosphate carrier deficiency |
| SLC25A38 | Anemia, sideroblastic, type 2 pyridoxine-refractory |
| SLC25A4 | Mitochondrial DNA depletion syndrome, type 12B (cardiomyopathic type) AR |
| SLC25A46 | Neuropathy, hereditary motor and sensory, type VIB |
| SLC26A2 | Achondrogenesis, type 1B (diastrophic dysplasia) |
| SLC26A3 | Diarrhea 1 secretory chloride, congenital |
| SLC26A4 | Deafness, autosomal recessive, type 4; Pendred syndrome |
| SLC26A5 | Deafness, autosomal recessive, type 61 |
| SLC27A4 | Ichthyosis prematurity syndrome |
| SLC29A3 | Histiocytosis-lymphadenopathy plus syndrome |
| SLC2A1 | GLUT1 deficiency syndrome 1, infantile onset, severe |
| SLC2A10 | Arterial tortuosity syndrome |
| SLC2A2 | Fanconi-Bickel syndrome |
| SLC2A9 | Hypouricemia, renal, type 2 |
| SLC30A10 | Hypermanganesemia with dystonia, type 1 |
| SLC33A1 | Congenital cataracts, hearing loss, and neurodegeneration |
| SLC34A1 | Hypercalcemia, infantile, type 2 |
| SLC34A2 | Pulmonary alveolar microlithiasis |
| SLC34A3 | Hypophosphatemic rickets with hypercalciuria |
| SLC35A1 | Congenital disorder of glycosylation, type 2F |
| SLC35A3 | Arthrogryposis, impaired intellectual development, and seizures |
| SLC35C1 | Congenital disorder of glycosylation, type 2C |
| SLC35D1 | Schneckenbecken dysplasia |
| SLC37A4 | Glycogen storage disease, type 1B |
| SLC38A8 | Foveal hypoplasia 2 with or without optic nerve misrouting and/or anterior segment dysgenesis |
| SLC39A13 | Ehlers-Danlos syndrome, spondylodysplastic type, 3 |
| SLC39A14 | Hypermanganesemia with dystonia 2 |
| SLC39A4 | Acrodermatitis enteropathica |
| SLC39A8 | Congenital disorder of glycosylation, type IIn |
| SLC3A1 | Cystinuria |
| SLC45A1 | Intellectual developmental disorder with neuropsychiatric features |
| SLC45A2 | Albinism, oculocutaneous, type 4 |
| SLC46A1 | Folate malabsorption, hereditary |
| SLC4A1 | Distal renal tubular acidosis |
| SLC4A11 | Corneal endothelial dystrophy, autosomal recessive |
| SLC4A4 | Renal tubular acidosis, proximal, with ocular abnormalities |
| SLC52A2 | Brown-Vialetto-Van Laere syndrome, type 2 |
| SLC52A3 | Brown-Vialetto-Van Laere syndrome, type 1 |
| SLC5A1 | Glucose/galactose malabsorption |
| SLC5A2 | Renal glucosuria |
| SLC5A5 | Thyroid dyshormonogenesis, type 1 |
| SLC5A7 | Myasthenic syndrome, congenital, type 20 presynaptic |
| SLC6A17 | intellectual developmental disorder, autosomal recessive 48 |
| SLC6A19 | Hartnup disorder |
| SLC6A3 | Parkinsonism-dystonia, infantile |
| SLC6A5 | Hyperekplexia, type 3 |
| SLC6A8 | Cerebral creatine deficiency syndrome, type 1 |
| SLC6A9 | Glycine encephalopathy with normal serum glycine |
| SLC7A14 | Retinitis pigmentosa 68 |
| SLC7A7 | Lysinuric protein intolerance |
| SLC7A9 | Cystinuria |
| SLC9A3 | Diarrhea 8 secretory sodium, congenital |
| SLC9A6 | Intellectual developmental disorder, X-linked syndromic, Christianson type |
| SLCO2A1 | Hypertrophic osteoarthropathy, primary, autosomal recessive, type 2 |
| SLITRK6 | Deafness and myopia |
| SLURP1 | Meleda disease |
| SLX4 | Fanconi anemia, complementation group P |
| SMARCAL1 | Schimke immunoosseous dysplasia |
| SMARCD2 | Specific granule deficiency 2 |
| SMG9 | Heart and brain malformation syndrome |
| SMN1 | Spinal muscular atrophy |
| SMN2 | {Spinal muscular atrophy, type III, modifier of} |
| SMOC1 | Microphthalmia. with limb anomalies |
| SMOC2 | Dentin dysplasia, type 1 with microdontia and misshapen teeth |
| SMPD1 | Niemann-Pick disease, type A; Niemann-Pick disease, type B |
| SMS | Intellectual developmental disorder, X-linked syndromic, Snyder-Robinson type |
| SNAP29 | Cerebral dysgenesis, neuropathy, ichthyosis, and palmoplantar keratoderma syndrome |
| SNX10 | Osteopetrosis, autosomal recessive, type 8 |
| SNX14 | Spinocerebellar ataxia, autosomal recessive, type 20 |
| SOBP | Impaired intellectual development, anterior maxillary protrusion, and strabismus |
| SOD1 | Spastic tetraplegia and axial hypotonia, progressive; Amyotrophic lateral sclerosis, type 1 |
| SOHLH1 | Ovarian dysgenesis 5 |
| SOST | Sclerosteosis, type 1; Van Buchem disease |
| SOX18 | Hypotrichosis-lymphedema-telangiectasia syndrome |
| SOX3 | Panhypopituitarism, X-linked |
| SP110 | Hepatic venoocclusive disease with immunodeficiency |
| SP7 | Osteogenesis imperfecta, type XII |
| SPAG1 | Ciliary dyskinesia, primary, type 28 |
| SPARC | Osteogenesis imperfecta, type XVII |
| SPART | Spactic paraplegia, type 20 autosomal recessive |
| SPATA5 | Neurodevelopmental disorder with hearing loss, seizures, and brain abnormalities |
| SPATA7 | Leber congenital amaurosis, type 3 |
| SPEG | Centronuclear myopathy, type 5 |
| SPG11 | Amyotrophic lateral sclerosis, type 5 juvenile |
| SPG21 | Mast syndrome |
| SPG7 | Spastic paraplegia, type 7 autosomal recessive |
| SPINK1 | Tropical calcific pancreatitis |
| SPINK5 | Netherton syndrome |
| SPINT2 | Diarrhea 3 secretory sodium, congenital, syndromic |
| SPR | Dystonia, dopa-responsive, due to sepiapterin reductase deficiency |
| SPRTN | Ruijs-Aalfs syndrome |
| SPTA1 | Pyropoikilocytosis; Apherocytosis, type 3 |
| SPTBN2 | Spinocerebellar ataxia, autosomal recessive, type 14 |
| SPTBN4 | Neurodevelopmental disorder with hypotonia, neuropathy, and deafness |
| SQSTM1 | Neurodegeneration with ataxia, dystonia, and gaze palsy, childhood-onset |
| SRD5A2 | 46,XY disorder of sex development due to 5-alpha-reductase 2 deficiency (pseudovaginal perineoscrotal hypospadias) |
| SRD5A3 | Congenital disorder of glycosylation, type 1Q; Kahrizi syndrome |
| SRPX2 | Rolandic epilepsy, impaired intellectual development, and speech dyspraxia |
| ST14 | Ichthyosis, congenital, autosomal recessive, type 11 |
| ST3GAL3 | Intellectual developmental disorder, autosomal recessive 12; Developmental and epileptic encephalopathy 15 |
| ST3GAL5 | Salt and pepper developmental regression syndrome |
| STAC3 | Native American myopathy |
| STAG3 | Premature ovarian failure 8 |
| STAMBP | Microcephaly-capillary malformation syndrome |
| STAR | Lipoid adrenal hyperplasia |
| STAT1 | Immunodeficiency, type 31B, mycobacterial and viral infections |
| STAT2 | Immunodeficiency, type 44 |
| STAT5B | Laron syndrome with immunodeficiency |
| STIL | Microcephaly, type 7 primary, autosomal recessive |
| STIM1 | Immunodeficiency, type 10 |
| STK4 | T-cell immunodeficiency, recurrent infections, autoimmunity, and cardiac malformations |
| STRA6 | Microphthalmia, isolated, with coloboma, type 8 |
| STRADA | Polyhydramnios, megalencephaly, and symptomatic epilepsy |
| STRC | Deafness, autosomal recessive, type 16 |
| STUB1 | Spinocerebellar ataxia, autosomal recessive, type 16 |
| STX11 | Hemophagocytic lymphohistiocytosis, familial, type 4 |
| STXBP2 | Hemophagocytic lymphohistiocytosis, familial, type 5 |
| SUCLA2 | Mitochondrial DNA depletion syndrome, type 5 (encephalomyopathic with or without methylmalonic aciduria) |
| SUCLG1 | Mitochondrial DNA depletion syndrome, type 9 (encephalomyopathic, type with methylmalonic aciduria) |
| SUFU | Joubert syndrome, type 32 |
| SUGCT | Glutaric aciduria, type 3 |
| SULT2B1 | Ichthyosis, congenital, autosomal recessive, type 14 |
| SUMF1 | Multiple sulfatase deficiency |
| SUN5 | Spermatogenic failure, type 16 |
| SUOX | Sulfite oxidase deficiency |
| SURF1 | Charcot-Marie-Tooth disease, type 4K; Leigh syndrome, due to COX IV deficiency |
| SYN1 | Epilepsy, X-linked, with variable learning disabilities and behavior disorders |
| SYNE1 | Spinocerebellar ataxia, autosomal recessive, type 8 |
| SYNE4 | Deafness, autosomal recessive, type 76 |
| SYNJ1 | Epileptic encephalopathy, early infantile, 53 |
| SYP | Intellectual developmental disorder, X-linked 96 |
| SYT14 | Spinocerebellar ataxia, autosomal recessive, type 11 |
| SZT2 | Epileptic encephalopathy, early infantile, 18 |
| TAC3 | Hypogonadotropic hypogonadism, type 10 with or without anosmia |
| TACO1 | Mitochondrial complex IV deficiency, nuclear type 8 |
| TACR3 | Hypogonadotropic hypogonadism, type 11 with or without anosmia |
| TACSTD2 | Corneal dystrophy, gelatinous drop-like |
| TAF1 | Dystonia-Parkinsonism, X-linked |
| TAF13 | Intellectual developmental disorder, autosomal recessive 60 |
| TAF2 | Intellectual developmental disorder, autosomal recessive 40 |
| TAF6 | Alazami-Yuan syndrome |
| TALDO1 | Transaldolase deficiency |
| TANGO2 | Metabolic encephalomyopathic crises, recurrent, with rhabdomyolysis, cardiac arrhythmias, and neurodegeneration |
| TAP1 | Bare lymphocyte syndrome, type 1 |
| TAP2 | Bare lymphocyte syndrome, type 1 due to TAP2 deficiency |
| TAPBP | Bare lymphocyte syndrome, type 1 |
| TAPT1 | Osteochondrodysplasia, complex lethal, Symoens-Barnes-Gistelinck type |
| TAT | Tyrosinemia, type 2 |
| TAZ | Barth syndrome |
| TBC1D20 | Warburg micro syndrome 4 |
| TBC1D23 | Pontocerebellar hypoplasia, type 11 |
| TBC1D24 | DOORS (deafness, onychodystrophy, osteodystrophy, impaired intellectual development, and seizures) syndrome; Epileptic encephalopathy, early infantile, type 16; Deafness, autosomal recessive, type 86 |
| TBC1D7 | Macrocephaly/megalencephaly syndrome, autosomal recessive |
| TBCD | Encephalopathy, progressive, early-onset, with brain atrophy and thin corpus callosum |
| TBCE | Encephalopathy, progressive, with amyotrophy and optic atrophy; Hypoparathyroidism-retardation-dysmorphism syndrome; Kenny-Caffey syndrome, type 1 |
| TBCK | Hypotonia, infantile, with psychomotor retardation and characteristic facies 3 |
| TBX15 | Cousin syndrome |
| TBX19 | Congenital isolated adrenocorticotropic hormone deficiency |
| TBXAS1 | Ghosal syndrome |
| TCAP | Limb-girdle muscular dystrophy, type 7 (LGMD R7) |
| TCIRG1 | Osteopetrosis, autosomal recessive, type 1 |
| TCN2 | Transcobalamin II deficiency |
| TCTN1 | Joubert syndrome, type 13 |
| TCTN2 | Joubert syndrome, type 24; ?Meckel syndrome, type 8 |
| TCTN3 | Joubert syndrome 18 |
| TDP1 | Spinocerebellar ataxia, autosomal recessive with axonal neuropathy |
| TDP2 | Spinocerebellar ataxia, autosomal recessive, type 23 |
| TDRD7 | Cataract 36 |
| TECPR2 | Spastic paraplegia, type 49 autosomal recessive |
| TECR | intellectual developmental disorder, autosomal recessive, type 14 |
| TECRL | Ventricular tachycardia, catecholaminergic polymorphic, 3 |
| TECTA | Deafness, autosomal recessive, type 21 |
| TELO2 | You-Hoover-Fong syndrome |
| TENM3 | Microphthalmia, isolated, with coloboma 9 |
| TERT | Dyskeratosis congenita, autosomal recessive, type 4 |
| TEX15 | Spermatogenic failure, type 25 |
| TF | Atransferrinemia |
| TFR2 | Hemochromatosis, type 3 |
| TFRC | Immunodeficiency, type 46 |
| TG | Thyroid dyshormonogenesis, type 3 |
| TGDS | Catel-Manzke syndrome |
| TGM1 | Ichthyosis, congenital, autosomal recessive, type 1 |
| TGM5 | Peeling skin syndrome, type 2 |
| TH | Segawa syndrome, recessive |
| THOC2 | Intellectual developmental disorder, X-linked 12 |
| THOC6 | Beaulieu-Boycott-Innes syndrome |
| THRB | Thyroid hormone resistance, autosomal recessive |
| TIMM50 | 3-methylglutaconic aciduria, type 9 |
| TIMM8A | Mohr-Tranebjaerg syndrome |
| TIMMDC1 | Mitochondrial complex I deficiency, nuclear type 31 |
| TJP2 | Cholestasis, progressive familial intrahepatic, type 4 |
| TK2 | Mitochondrial DNA depletion syndrome , type 2 (myopathic type) |
| TKT | Short stature, developmental delay, and congenital heart defects |
| TLE6 | Preimplantation embryonic lethality |
| TMC1 | Deafness, autosomal recessive, type 7 |
| TMC6 | Epidermodysplasia verruciformis |
| TMC8 | Epidermodysplasia verruciformis |
| TMCO1 | Craniofacial dysmorphism, skeletal anomalies, and impaired intellectual development 1 |
| TMEM107 | Meckel syndrome, type 13; Orofaciodigital syndrome, type 16 |
| TMEM126A | Optic atrophy 7 |
| TMEM126B | Mitochondrial complex I deficiency, nuclear type 29 |
| TMEM138 | Joubert syndrome 16 |
| TMEM165 | Congenital disorder of glycosylation, type 2K |
| TMEM199 | Congenital disorder of glycosylation, type 2P |
| TMEM216 | Joubert syndrome, type 2; Meckel syndrome, type 2 |
| TMEM231 | Joubert syndrome, type 20; Meckel syndrome,type 11 |
| TMEM237 | Joubert syndrome, type 14 |
| TMEM260 | Structural heart defects and renal anomalies syndrome |
| TMEM38B | Osteogenesis imperfecta, type XIV |
| TMEM67 | Joubert syndrome, type 6; Meckel syndrome, type 3; COACH syndrome |
| TMEM70 | Mitochondrial complex V (ATP synthase) deficiency, nuclear type 2 |
| TMIE | Deafness, autosomal recessive, type 6 |
| TMPRSS15 | Enterokinase deficiency |
| TMPRSS3 | Deafness, autosomal recessive, type 45573 |
| TMPRSS6 | Iron-refractory iron deficiency anemia |
| TMTC3 | Lissencephaly 8 |
| TNFRSF11A | Osteopetrosis, autosomal recessive, type 7 |
| TNFRSF11B | Paget disease of bone, type 5 juvenile-onset |
| TNFRSF13B | Immunodeficiency, common variable, type 2 |
| TNFSF11 | Osteopetrosis, autosomal recessive, type 2 |
| TNIK | Intellectual developmental disorder, autosomal recessive 54 |
| TNNT1 | Nemaline myopathy , type 5 Amish type |
| TNXB | Ehlers-Danlos syndrome, classic-like |
| TOE1 | Pontocerebellar hypoplasia, type 7 |
| TOP3A | Microcephaly, growth restriction, and increased sister chromatid exchange 2 |
| TP53RK | Galloway-Mowat syndrome 4 |
| TPI1 | Hemolytic anemia due to triosephosphate isomerase deficiency |
| TPK1 | Episodic encephalopathy due to thiamine pyrophosphokinase deficiency |
| TPM3 | Nemaline myopathy, type 1; Congenital fiber-type disproportion myopathy |
| TPO | Thyroid dyshormonogenesis, type 2A |
| TPP1 | Ceroid lipofuscinosis, neuronal, type 2; Spinocerebellar ataxia, autosomal recessive, type 7 |
| TPRN | Deafness, autosomal recessive, type 79 |
| TRAF3IP1 | Senior-Loken syndrome, type 9 |
| TRAIP | Seckel syndrome, type 9 |
| TRAPPC11 | Limb-girdle muscular dystrophy, type 18 (LGMD R18) |
| TRAPPC12 | Encephalopathy, progressive, early-onset, with brain atrophy and spasticity |
| TRAPPC6B | Neurodevelopmental disorder with microcephaly, epilepsy, and brain atrophy |
| TRAPPC9 | Intellectual developmental disorder, autosomal recessive 13 |
| TRDN | Ventricular tachycardia, catecholaminergic polymorphic, type 5 with or without muscle weakness |
| TREM2 | Nasu-Hakola disease |
| TREX1 | Aicardi-Goutieres syndrome, type 1 |
| TRHR | Hypothyroidism, congenital, nongoitrous, type 7 |
| TRIM2 | Charcot-Marie-Tooth disease, type 2R |
| TRIM32 | Limb-girdle muscular dystrophy, type 8 (LGMD R8) |
| TRIM37 | Mulibrey nanism |
| TRIOBP | Deafness, autosomal recessive, type 28 |
| TRIP11 | Achondrogenesis, type 1A |
| TRIP13 | Mosaic variegated aneuploidy syndrome 3 |
| TRIP4 | Spinal muscular atrophy with congenital bone fractures 1 |
| TRIT1 | Combined oxidative phosphorylation deficiency 35 |
| TRMT10A | Microcephaly, short stature, and impaired glucose metabolism 1 |
| TRMT10C | Combined oxidative phosphorylation deficiency 30 |
| TRMT5 | Combined oxidative phosphorylation deficiency 26 |
| TRMU | Liver failure, transient infantile |
| TRNT1 | Retinitis pigmentosa and erythrocytic microcytosis |
| TRPM1 | Night blindness, congenital stationary (complete), type 1C, autosomal recessive |
| TRPM6 | Familial hypomagnesemia with secondary hypocalcemia |
| TRPV6 | Hyperparathyroidism, transient neonatal |
| TSEN15 | Pontocerebellar hypoplasia, type 2F |
| TSEN2 | Pontocerebellar hypoplasia, type 2B |
| TSEN34 | Pontocerebellar hypoplasia type 2C |
| TSEN54 | Pontocerebellar hypoplasia, type 2A; Pontocerebellar hypoplasia, type 4 |
| TSFM | Combined oxidative phosphorylation deficiency, type 3 |
| TSHB | Hypothyroidism, congenital, nongoitrous, type 4 |
| TSHR | Hypothyroidism, congenital, nongoitrous, type 1 |
| TSPAN7 | Intellectual developmental disorder, X-linked 58 |
| TSPYL1 | Sudden infant death with dysgenesis of the testes syndrome |
| TTC19 | Mitochondrial complex III deficiency, nuclear type 2 |
| TTC21B | Short-rib thoracic dysplasia, type 4 with or without polydactyly |
| TTC37 | Trichohepatoenteric syndrome 1 |
| TTC7A | Gastrointestinal defects and immunodeficiency syndrome |
| TTC8 | Bardet-Biedl syndrome, type 8 |
| TTI2 | Intellectual developmental disorder, autosomal recessive 39 |
| TTLL5 | Cone-rod dystrophy 19 |
| TTN | Limb-girdle muscular dystrophy type 10 (LGMDR10); Early-onset myopathy with fatal cardiomyopathy (Salih myopathy) |
| TTPA | Ataxia with isolated vitamin E deficiency |
| TUBA8 | Cortical dysplasia, complex, with other brain malformations, type 8 |
| TUBGCP4 | Microcephaly and chorioretinopathy, autosomal recessive, type 3 |
| TUBGCP6 | Microcephaly and chorioretinopathy, autosomal recessive, type 1 |
| TUFM | Combined oxidative phosphorylation deficiency 4 |
| TULP1 | Leber congenital amaurosis, type 15 |
| TUSC3 | Intellectual developmental disorder, autosomal recessive 7 |
| TWIST2 | Focal facial dermal dysplasia, type 3 (Setleis type) |
| TWNK | Mitochondrial DNA depletion syndrome, type 7 (hepatocerebral type); Perrault syndrome type 5 |
| TXNL4A | Burn-McKeown syndrome |
| TYK2 | Immunodeficiency, type 35 |
| TYMP | Mitochondrial DNA depletion syndrome, type 1 (MNGIE type) |
| TYR | Oculocutaneous albinism (OCA) type 1A; OCA type 1B |
| TYROBP | Polycystic lipomembranous osteodysplasia with sclerosing leukoencephalopathy, type 1 (Nasu-Hakola disease) |
| TYRP1 | Albinism, oculocutaneous, type 3 |
| UBA1 | Spinal muscular atrophy, X-linked 2, infantile |
| UBA5 | Epileptic encephalopathy, early infantile, 44 |
| UBE2A | Intellectual developmental disorder, X-linked syndromic, Nascimento type |
| UBE2T | Fanconi anemia, complementation group T |
| UBE3A | Angelman syndrome |
| UBE3B | Kaufman oculocerebrofacial syndrome |
| UBR1 | Johanson-Blizzard syndrome |
| UCHL1 | Spastic paraplegia, type 79 autosomal recessive |
| UFM1 | Leukodystrophy, hypomyelinating, type 14 |
| UGT1A1 | Crigler-Najjar syndrome, type 1; Crigler-Najjar syndrome, type 2 |
| UMPS | Orotic aciduria |
| UNC13D | Hemophagocytic lymphohistiocytosis, familial, type 3 |
| UNC80 | Hypotonia, infantile, with psychomotor retardation and characteristic facies, type 2 |
| UNG | Immunodeficiency with hyper IgM, type 5 |
| UPB1 | Beta-ureidopropionase deficiency |
| UPF3B | Intellectual developmental disorder, X-linked syndromic 14 |
| UQCRB | Mitochondrial complex III deficiency, nuclear, type 3 |
| UQCRC2 | Mitochondrial complex III deficiency, nuclear type 5 |
| UQCRQ | Mitochondrial complex III deficiency, nuclear, type 4 |
| UROD | Porphyria cutanea tarda |
| UROS | Porphyria, congenital erythropoietic |
| USB1 | Poikiloderma with neutropenia |
| USH1C | Usher syndrome, type 1C; Deafness, autosomal recessive, type 18A |
| USH1G | Usher syndrome, type 1G |
| USH2A | Usher syndrome, type 2A |
| USP18 | Pseudo-TORCH syndrome 2 |
| USP9X | Intellectual developmental disorder, X-linked 99 |
| UVSSA | UV-sensitive syndrome, type 3 |
| VAC14 | Striatonigral degeneration, childhood-onset |
| VARS1 | Neurodevelopmental disorder with microcephaly, seizures, and cortical atrophy |
| VARS2 | Combined oxidative phosphorylation deficiency 20 |
| VDR | Rickets, vitamin D-resistant, type 2A |
| VIPAS39 | Arthrogryposis, renal dysfunction and cholestasis, type 2 |
| VKORC1 | Vitamin K-dependent clotting factors, combined deficiency of, type 2 |
| VLDLR | Cerebellar hypoplasia, impaired intellectual development, and dysequilibrium syndrome 1 |
| VPS13A | Choreoacanthocytosis |
| VPS13B | Cohen syndrome |
| VPS13C | Parkinson disease 23 autosomal recessive, early onset |
| VPS33B | Arthrogryposis, renal dysfunction and cholestasis, type 1 |
| VPS37A | Spastic paraplegia, type 53 autosomal recessive |
| VPS45 | Neutropenia, severe congenital, type 5 |
| VPS53 | Pontocerebellar hypoplasia, type 2E |
| VRK1 | Pontocerebellar hypoplasia, type 1A |
| VSX2 | Microphthalmia with coloboma 3; Isolated microphthalmia 2 |
| VWF | von Willibrand disease, type 3 |
| WARS2 | Neurodevelopmental disorder, mitochondrial, with abnormal movements and lactic acidosis, with or without seizures |
| WAS | Wiskott-Aldrich syndrome; Thrombocytopenia, X-linked |
| WASHC4 | Intellectual developmental disorder, autosomal recessive 43 |
| WASHC5 | Ritscher-Schinzel syndrome, type 1 |
| WDR19 | Nephronophthisis, type 13; Senior-Loken syndrome, type 8 |
| WDR35 | Cranioectodermal dysplasia 2 |
| WDR45B | Neurodevelopmental disorder with spastic quadriplegia and brain abnormalities with or without seizures |
| WDR62 | Microcephaly, type 2 primary, autosomal recessive, with or without cortical malformations |
| WDR72 | Amelogenesis imperfecta, type 2A3 (hypomaturation type) |
| WDR73 | Galloway-Mowat syndrome 1 |
| WDR81 | Cerebellar ataxia, impaired intellectual development, and dysequilibrium syndrome, type 2; Hydrocephalus, congenital, 3, with brain anomalies |
| WEE2 | Oocyte maturation defect 5 |
| WFS1 | Wolfram syndrome, type 1 |
| WHRN | Usher syndrome, type 2D; Deafness, autosomal recessive, type 31 |
| WIPF1 | Wiskott-Aldrich syndrome 2 |
| WNK1 | Neuropathy, hereditary sensory and autonomic, type 2 |
| WNT1 | Osteogenesis imperfecta, type XV |
| WNT10A | Ectodermal dysplasia 16 (odontoonychodermal dysplasia) |
| WNT10B | Split-hand/foot malformation, type 6 |
| WNT3 | Tetra-amelia syndrome |
| WNT7A | Fuhrmann syndrome |
| WRAP53 | Dyskeratosis congenita, autosomal recessive, type 3 |
| WRN | Werner syndrome |
| WWOX | Epileptic encephalopathy, early infantile, type 28; Spinocerebellar ataxia, autosomal recessive, type 12 |
| XDH | Xanthinuria, type 1 |
| XIAP | Lymphoproliferative syndrome, X-linked, 2 |
| XPA | Xeroderma pigmentosum, group A |
| XPC | Xeroderma pigmentosum, group C |
| XPNPEP3 | Nephronophthisis-like nephropathy, type 1 |
| XRCC4 | Short stature, microcephaly, and endocrine dysfunction |
| XYLT1 | Desbuquois dysplasia, type 2 |
| XYLT2 | Spondyloocular syndrome |
| YARS2 | Myopathy, lactic acidosis, and sideroblastic anemia, type 2 |
| YY1AP1 | Grange syndrome |
| ZAP70 | Autoimmune disease, multisystem, infantile-onset, type 2; Immunodeficiency, type 48 |
| ZBTB24 | Immunodeficiency-centromeric instability-facial anomalies syndrome, type 2 |
| ZC3H14 | intellectual developmental disorder, autosomal recessive, type 56 |
| ZDHHC9 | Intellectual developmental disorder, X-linked syndromic, Raymond type |
| ZFYVE26 | Spastic paraplegia, type 15 autosomal recessive |
| ZMPSTE24 | Mandibuloacral dysplasia with, type B lipodystrophy |
| ZMYND10 | Ciliary dyskinesia, primary, type 22 |
| ZNF408 | Retinitis pigmentosa, type 72 |
| ZNF423 | Joubert syndrome, type 19 |
| ZNF469 | Brittle cornea syndrome, type 1 |
| ZNF711 | Intellectual developmental disorder, X-linked 97 |
| ZNHIT3 | PEHO syndrome |
| ZP1 | Oocyte maturation defect, type 1 |
Список заболеваний, которые анализирует
PLUS
| ACADM | Medium-chain acyl-CoA dehydrogenase deficiency |
| AGXT | Hyperoxaluria, primary, type 1 |
| ARSA | Metachromatic leukodystrophy |
| ATP7B | Wilson disease |
| BTD | Biotinidase deficiency |
| CBS | Homocystinuria due to cystathionine beta-synthase |
| CFTR | Cystic fibrosis |
| DHCR7 | Smith-Lemli-Opitz syndrome |
| EMD | Emery-Dreifuss muscular dystrophy, type 1 X-linked |
| FMR1 | Fragile X syndrome |
| GAA | Glycogen storage disease, type 2 |
| GALC | Krabbe disease |
| GALT | Galactosemia |
| GBA | Gaucher Disease |
| GJB1 | Charcot-Marie-Tooth neuropathy, X-linked dominant, type 1 |
| GJB2 | Deafness, autosomal recessive, type 1A; Deafness, digenic, GJB2/GJB6 |
| GJB6 | Deafness, autosomal recessive, type 1B; Deafness, digenic GJB2/GJB6 |
| GLA | Fabry disease |
| HADHA | Long-chain 3-hydroxyl-CoA dehydrogenase (LCHAD) deficiency; Mitochondrial trifunctional protein deficiency |
| HBA1 | Alpha-thalassemia |
| HBA2 | Alpha-thalassemia |
| HBB | Beta-thalassemia, Sickle hemoglobinopathies cell anemia and other HBB-related |
| HEXA | Tay-Sachs disease |
| MEFV | Familial Mediterranean fever |
| MMACHC | Methylmalonic aciduria and homocystinuria, cblC type |
| PAH | Phenylketonuria |
| PMM2 | Congenital disorder of glycosylation, type 1A |
| SERPINA1 | Alpha-1 antitrypsin deficiency |
| SLC26A2 | Achondrogenesis, type 1B (diastrophic dysplasia) |
| SMN1 | Spinal muscular atrophy |
Список заболеваний, которые анализирует
ADVANCE
| AAAS | Triple-A syndrome (achalasia-addisonianism-alacrimia) |
| ABAT | GABA-transaminase deficiency |
| ABCA12 | Ichthyosis congenital autosomal recessive type 4A and 4B (harlequin) |
| ABCA3 | Surfactant metabolism dysfunction, pulmonary, type 3 |
| ABCC8 | Hyperinsulinemic hypoglycemia, type 1 (congenital hyperinsulinism); Permanent neonatal diabetes mellitus (PNDM) |
| ABCD1 | Adrenoleukodystrophy |
| ABHD12 | PHARC syndrome (polyneuropathy, hearing loss, ataxia, retinitis pigmentosa and cataract) |
| ACAD9 | Acyl-CoA dehydrogenase 9 deficiency (mitochondrial complex I deficiency, nuclear, type 20) |
| ACADM | Medium-chain acyl-CoA dehydrogenase deficiency |
| ACADS | Short-chain acyl-CoA dehydrogenase deficiency |
| ACADVL | Very long-chain acyl-CoA dehydrogenase (VLCAD) deficiency |
| ACAT1 | Alpha-methylacetoacetic aciduria (3-ketothiolase deficiency) |
| ACE | Renal tubular dysgenesis |
| ACOX1 | Peroxisomal acyl-CoA oxidase deficiency |
| ACTA1 | Nemaline myopathy 3; Congenital fiber-type disproportion myopathy 1 |
| ADA | Severe combined immunodeficiency due to adenosine deaminase deficiency (ADA) |
| ADAMTS10 | Weill-Marchesani syndrome, type 1 recessive |
| ADAMTS13 | Thrombotic thrombocytopenic purpura, familial (Schulman-Upshaw syndrome) |
| AFF2 | Intellectual developmental disorder, X-linked 109 |
| AGA | Aspartylglucosaminuria (glycosylasparaginase deficiency) |
| AGL | Glycogen storage disease, type 3 |
| AGPAT2 | Congenital generalized lipodystrophy (Berardinelli-Seip syndrome) |
| AGPS | Rhizomelic chondrodysplasia punctata, type 3 |
| AGRN | Myasthenic syndrome, congenital, type 8 |
| AGXT | Hyperoxaluria, primary, type 1 |
| AHI1 | Joubert syndrome, type 3 |
| AIMP1 | Leukodystrophy, hypomyelinating, type 3 |
| AIRE | Autoimmune polyendocrinopathy syndrome, type 1 |
| ALDH18A1 | Spastic paraplegia, type 9B, autosomal recessive; Cutis laxa, type 3A (De Barsy syndrome) |
| ALDH3A2 | Sjogren-Larsson syndrome |
| ALDH5A1 | Succinic semialdehyde dehydrogenase deficiency |
| ALDH7A1 | Epilepsy, pyridoxine-dependent |
| ALDOB | Fructose intolerance, hereditary |
| ALG12 | Congenital disorder of glycosylation, type 1G |
| ALG6 | Congenital disorder of glycosylation, type 1C |
| ALMS1 | Alström syndrome |
| ALOX12B | Ichthyosis, congenital, autosomal recessive, type 2 |
| ALPL | Hypophosphatasia, infantile/childhood |
| AMACR | Bile acid synthesis defect, congenital, type 4; Alpha-methylacyl-CoA racemase deficiency |
| AMT | Glycine encephalopathy |
| ANO10 | Spinocerebellar ataxia, autosomal recessive, type 10 |
| ANTXR2 | Hyaline fibromatosis syndrome |
| AP1S2 | Pettigrew syndrome |
| APTX | Ataxia, early-onset, with oculomotor apraxia and hypoalbuminemia |
| AQP2 | Diabetes insipidus, nephrogenic, type 2 |
| ARFGEF2 | Periventricular heterotopia with microcephaly |
| ARL13B | Joubert syndrome type 8 |
| ARL6 | Bardet-Biedl syndrome, type 3 |
| ARSA | Metachromatic leukodystrophy |
| ARSB | Mucopolysaccharidosis, type 6 (Maroteaux-Lamy syndrome) |
| ARSE | Chondrodysplasia punctata, X-linked recessive |
| ARX | Epileptic encephalopathy, early infantile, type 1; ARX-related developmental disorders |
| ASL | Argininosuccinic aciduria |
| ASPA | Canavan disease |
| ASPM | Primary microcephaly type 5 autosomal recessive |
| ASS1 | Citrullinemia, type 1 |
| ATM | Ataxia-telangiectasia |
| ATP6V0A2 | Cutis laxa, autosomal recessive, type 2A; Wrinkly skin syndrome |
| ATP7A | Menkes disease; Occipital horn syndrome |
| ATP7B | Wilson disease |
| ATR | Seckel syndrome, type 1 |
| ATRX | Intellectual disability-hypotonic facies syndrome, X-linked; Alpha-thalassemia/intellectual developmental disorder syndrome |
| AUH | 3-methylglutaconic aciduria, type 1 |
| BBS1 | Bardet-Biedl syndrome, type 1 |
| BBS10 | Bardet-Biedl syndrome, type 10 |
| BBS12 | Bardet-Biedl syndrome, type 12 |
| BBS2 | Bardet-Biedl syndrome, type 2 |
| BBS7 | Bardet-Biedl syndrome, type 7 |
| BBS9 | Bardet-Biedl syndrome, type 9 |
| BCKDHA | Maple syrup urine disease, type 1A |
| BCKDHB | Maple syrup urine disease, type 1B |
| BCS1L | Mitochondrial complex III deficiency nuclear type 1; GRACILE syndrome; Bjornstad syndrome |
| BEST1 | Bestrophinopathy, AR |
| BLM | Bloom syndrome |
| BRWD3 | Intellectual developmental disorder, X-linked 93 |
| BSCL2 | Congenital generalized lipodystrophy, type 2; Encephalopathy, progressive, with or without lipodystrophy |
| BTD | Biotinidase deficiency |
| BUB1B | Mosaic variegated aneuploidy syndrome 1 |
| CBS | Homocystinuria due to cystathionine beta-synthase |
| CC2D2A | Joubert syndrome, type 9; Meckel syndrome, type 6; COACH syndrome, 2 |
| CCDC88C | Hydrocephalus, congenital, type 1 |
| CD40LG | Hyper-IgM syndrome, type 1 (immunodeficiency, X-linked, with hyper-IgM, type 1) |
| CDAN1 | Dyserythropoietic anemia, congenital, type 1A |
| CDH23 | Deafness, autosomal recessive, type 12; Usher syndrome, type 1D |
| CDK5RAP2 | Primary microcephaly type 3 autosomal recessive |
| CENPJ | Primary microcephaly type 6 autosomal recessive |
| CEP152 | Primary microcephaly type 9 autosomal recessive |
| CEP290 | Meckel syndrome, type 4; Joubert syndrome, type 5; Leber congenital amaurosis, type 10 |
| CFL2 | Nemaline myopathy, type 7 autosomal recessive |
| CFTR | Cystic fibrosis |
| CHAT | Myasthenic syndrome, congenital, type 6 presynaptic |
| CHM | Choroideremia |
| CHRND | Myasthenic syndrome, congenital, type 3B, fast-channel; Multiple pterygium syndrome, lethal type |
| CHRNE | Myasthenic syndrome, congenital, type 4B, fast-channel; Myasthenic syndrome, congenital, type 4C, associated with acetylcholine receptor deficiency |
| CHRNG | Multiple pterygium syndrome (MPS), Escobar type; MPS, lethal type |
| CHST14 | Ehlers-Danlos syndrome, musculocontractural, type 1 |
| CHST3 | Spondyloepiphyseal dysplasia with congenital joint dislocations |
| CLCN1 | Myotonia congenita, recessive |
| CLN3 | Ceroid lipofuscinosis, neuronal, type 3 |
| CLN5 | Ceroid lipofuscinosis, neuronal, type 5 |
| CLN6 | Ceroid lipofuscinosis, neuronal, type 6 |
| CLN8 | Ceroid lipofuscinosis, neuronal, type 8 |
| CLRN1 | Usher syndrome, type 3A |
| CNGB3 | Achromatopsia, type 3 |
| COG4 | Congenital disorder of glycosylation, type 2J |
| COL11A2 | Otospondylomegaepiphyseal dysplasia, autosomal recessive |
| COL17A1 | Epidermolysis bullosa, junctional, non-Herlitz type |
| COL4A3 | Alport syndrome, autosomal recessive, type 2 |
| COL4A5 | Alport syndrome, X-linked |
| COL7A1 | Dystrophic epidermolysis bullosa (DEB), Hallopeau-Siemens (HS) type and non-HS type; DEB pruriginosa; DEB pretibial |
| COLQ | Myasthenic syndrome, congenital, type 5 |
| COQ2 | Primary coenzyme Q10 deficiency, type 1 |
| CORO1A | Immunodeficiency, type 8 |
| CPS1 | Carbamoylphosphate synthetase 1 deficiency |
| CPT1A | Carnitine palmitoyltransferase type 1A deficiency, hepatic |
| CPT2 | Carnitine palmitoyltransferase type 2 deficiency, lethal neonatal; Carnitine palmitoyltransferase type 2 deficiency, infantile |
| CRLF1 | Cold-induced sweating syndrome type 1 |
| CRTAP | Osteogenesis imperfecta, type 7 |
| CSTB | Epilepsy, progressive myoclonic type 1A (Unverricht and Lundborg) |
| CTC1 | Cerebroretinal microangiopathy with calcifications and cysts |
| CTNS | Nephropathic cystinosis |
| CTSD | Ceroid lipofuscinosis, neuronal, type 10 |
| CTSK | Pycnodysostosis |
| CUL4B | Intellectual developmental disorder, X-linked syndromic, Cabezas type |
| CUL7 | 3M syndrome 1 |
| CYBB | Chronic granulomatous disease, X-linked |
| CYP11A1 | 46,XY disorder of sex development-adrenal insufficiency due to CYP11A1 deficiency |
| CYP1B1 | Glaucoma, primary congenital, type 3A |
| CYP21A2 | Congenital adrenal hyperplasia due to 21-hydroxylase deficiency |
| CYP27A1 | Cerebrotendinous xanthomatosis |
| CYP27B1 | Vitamin D-dependent rickets, type 1 |
| D2HGDH | D-2-hydroxyglutaric aciduria |
| DBH | Dopamine beta-hydroxylase deficiency |
| DBT | Maple syrup urine disease, type 2 |
| DCLRE1C | Omenn syndrome; Severe combined immunodeficiency, Athabascan type |
| DCX | Lissencephaly, X-linked, type 1 |
| DDC | Aromatic L-amino acid decarboxylase deficiency |
| DGUOK | DGUOK-related mitochondrial DNA depletion syndrome |
| DHCR7 | Smith-Lemli-Opitz syndrome |
| DHDDS | Retinitis pigmentosa, type 59 |
| DHH | 46,XY complete gonadal dysgenesis |
| DLAT | Pyruvate dehydrogenase E2 deficiency |
| DLG3 | Intellectual developmental disorder, X-linked 90 |
| DMD | Duchenne/Becker muscular dystrophy |
| DMD | Duchenne/Becker muscular dystrophy |
| DMP1 | Hypophosphatemic rickets, autosomal recessive |
| DNAH11 | Ciliary dyskinesia, primary, type 7 with or without situs inversus |
| DNAH5 | Ciliary dyskinesia, primary, type 3 with or without situs inversus |
| DNAL1 | Ciliary dyskinesia, primary, type 16 |
| DOCK8 | Hyper-IgE recurrent infection syndrome, autosomal recessive |
| DOK7 | Fetal akinesia deformation sequence, type 3; Myasthenic syndrome, congenital, type 10 |
| DPAGT1 | Congenital disorder of glycosylation, type 1J; Myasthenic syndrome, congenital, type 13 |
| DPYD | Dihydropyrimidine dehydrogenase deficiency |
| DYNC2H1 | Short-rib thoracic dysplasia, type 3 with or without polydactyly |
| EDA | Ectodermal dysplasia, type 1 hypohidrotic, X-linked |
| EDNRB | ABCD syndrome |
| EIF2AK3 | Wolcott-Rallison syndrome |
| EIF2B2 | Leukoencephalopathy with vanishing white matter (VWM) |
| EIF2B3 | Leukoencephalopathy with vanishing white matter (VWM) |
| EIF2B4 | Leukoencephalopathy with vanishing white matter (VWM) |
| EIF2B5 | Leukoencephalopathy with vanishing white matter (VWM) |
| EMD | Emery-Dreifuss muscular dystrophy, type 1 X-linked |
| ENPP1 | Arterial calcification, generalized, of infancy, type 1 |
| EPG5 | Vici syndrome |
| EPM2A | Epilepsy, progressive myoclonic, type 2A (Lafora) |
| ERCC2 | Trichothiodystrophy, type 1 |
| ERCC4 | Fanconi anemia, complementation group Q |
| ERCC5 | Cerebrooculofacioskeletal syndrome, type 3 |
| ERCC6 | Cockayne syndrome, type B; Cerebrooculofacioskeletal syndrome, type 1 |
| ERCC8 | Cockayne syndrome, type A |
| ESCO2 | Roberts syndrome; Juberg-Hayward syndrome |
| ETFDH | Glutaric acidemia, type 2C |
| ETHE1 | Ethylmalonic encephalopathy |
| EVC | Ellis-van Creveld syndrome |
| EVC2 | Ellis-van Creveld syndrome |
| EXOSC3 | Pontocerebellar hypoplasia, type 1B |
| F11 | Factor XI deficiency |
| F8 | Hemophilia A |
| F9 | Hemophilia B |
| FA2H | Spastic paraplegia, type 35 autosomal recessive |
| FAH | Tyrosinemia, type 1 |
| FAM20C | Raine syndrome |
| FANCA | Fanconi anemia, complementation group A |
| FANCC | Fanconi anemia, complementation group C |
| FANCF | Fanconi anemia, complementation group F |
| FANCG | Fanconi anemia, complementation group G |
| FANCI | Fanconi anemia, complementation group I |
| FGD1 | Aarskog-Scott syndrome; intellectual developmental disorder, X-linked syndromic, type 16 |
| FKRP | Muscular dystrophy-dystroglycanopathy, type 5A (Walker-Warburg syndrome); Type 5B; Type 5C (limb-girdle muscular dystrophy, type 9 [LGMDR9]) |
| FKTN | Muscular dystrophy-dystroglycanopathy, type 4A (Walker-Warburg syndrome); Type 4B; Type 4C (limb-girdle muscular dystrophy, type 13 [LGMD R13]) |
| FMO3 | Trimethylaminuria |
| FMR1 | Fragile X syndrome |
| FRAS1 | Fraser syndrome, type 1 |
| FTSJ1 | Intellectual developmental disorder, X-linked 9 |
| FUCA1 | Fucosidosis |
| G6PC | Glycogen storage disease, type 1A |
| G6PD | Hemolytic anemia, G6PD deficient (favism) |
| GAA | Glycogen storage disease, type 2 |
| GALC | Krabbe disease |
| GALT | Galactosemia |
| GAMT | Cerebral creatine deficiency syndrome, type 2 |
| GAN | Giant axonal neuropathy, type 1 |
| GATM | Cerebral creatine deficiency syndrome, type 3 |
| GBA | Gaucher Disease |
| GBE1 | Glycogen storage disease, type 4 |
| GCDH | Glutaricaciduria, type 1 |
| GFM1 | Combined oxidative phosphorylation deficiency, type 1 |
| GFPT1 | Myasthenia, congenital, type 12 with tubular aggregates |
| GJB1 | Charcot-Marie-Tooth neuropathy, X-linked dominant, type 1 |
| GJB2 | Deafness, autosomal recessive, type 1A; Deafness, digenic, GJB2/GJB6 |
| GJB6 | Deafness, autosomal recessive, type 1B; Deafness, digenic GJB2/GJB6 |
| GLA | Fabry disease |
| GLB1 | GM1-gangliosidosis, types 1-3; Mucopolysaccharidosis, type 4B (Morquio) |
| GLDC | Glycine encephalopathy |
| GLRA1 | Hyperekplexia, type 1 |
| GNPTAB | Mucolipidosis 2 alpha/beta; Mucolipidosis 3 alpha/beta |
| GNRHR | Hypogonadotropic hypogonadism, type 7 without anosmia |
| GOSR2 | Epilepsy, progressive myoclonic, type 6 |
| GP1BA | Bernard-Soulier syndrome, type A1 |
| GP1BB | Bernard-Soulier syndrome, type B |
| GPHN | Molybdenum cofactor deficiency C |
| GPR143 | Ocular albinism, type 1 (Nettleship-Falls type) |
| GRHPR | Hyperoxaluria, primary, type 2 |
| GRIP1 | Fraser syndrome 3 |
| HADH | 3-hydroxyacyl-CoA dehydrogenase deficiency |
| HADHA | Long-chain 3-hydroxyl-CoA dehydrogenase (LCHAD) deficiency; Mitochondrial trifunctional protein deficiency |
| HBA1 | Alpha-thalassemia |
| HBA2 | Alpha-thalassemia |
| HBB | Beta-thalassemia, Sickle cell anemia and other HBB-related hemoglobinopathies |
| HEXA | Tay-Sachs disease |
| HEXB | Sandhoff disease, infantile, juvenile, and adult forms |
| HGSNAT | Mucopolysaccharidosis type 3C (Sanfilippo syndrome C) |
| HIBCH | 3-hydroxyisobutryl-CoA hydrolase deficiency |
| HLCS | Holocarboxylase synthetase deficiency |
| HPS3 | Hermansky-Pudlak syndrome, type 3 |
| HPS4 | Hermansky-Pudlak syndrome, type 4 |
| HPS6 | Hermansky-Pudlak syndrome, type 6 |
| HSD17B3 | 46,XY disorder of sex development due to 17-beta-hydroxysteroid dehydrogenase 3 deficiency |
| HSD17B4 | D-bifunctional protein deficiency |
| HSD3B2 | Adrenal hyperplasia, congenital, due to 3-beta-hydroxysteroid dehydrogenase 2 deficiency |
| HSPG2 | Dyssegmental dysplasia, Silverman-Handmaker type |
| IDS | Mucopolysaccharidosis, type 2 |
| IDUA | Mucopolysaccharidosis type 1 |
| IGHMBP2 | Charcot-Marie-Tooth disease, axonal, type 2S |
| IL1RAPL1 | Intellectual developmental disorder, X-linked 21 |
| IL2RG | Severe combined immunodeficiency, X-linked |
| IL7R | Severe combined immunodeficiency, T-cell negative, B-cell/natural killer cell-positive type |
| INPP5E | Joubert syndrome, type 1 |
| INSR | Diabetes mellitus, insulin-resistant, with acanthosis nigricans, type A |
| ITGB4 | Epidermolysis bullosa, junctional, with pyloric atresia |
| IVD | Isovaleric acidemia |
| JAK3 | Severe Combined Immunodeficiency, autosomal recessive, T-negative/B-positive type |
| KCNJ1 | Bartter syndrome, type 2 |
| KCNJ13 | Leber congenital amaurosis, type 16 |
| KCTD7 | Epilepsy, progressive myoclonic, type 3 with or without intracellular inclusions |
| KDM5C | Intellectual developmental disorder, X-linked syndromic, Claes-Jensen type |
| KIF7 | Acrocallosal syndrome; Joubert syndrome, type 12 |
| L1CAM | L1 Syndrome |
| LAMA2 | LAMA2-related muscular dystrophy |
| LAMA3 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LAMB2 | Pierson syndrome; Nephrotic syndrome, type 5 with or without ocular abnormalities |
| LAMB3 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LAMC2 | Junctional epidermolysis bullosa (JEB) Herlitz type; JEB non-Herlitz type |
| LIFR | Stuve-Wiedemann syndrome / Schwartz-Jampel type 2 syndrome |
| LMNA | Cardiomyopathy, dilated, 1A |
| LPL | Lipoprotein lipase deficiency |
| LRP2 | Donnai-Barrow syndrome |
| LRPPRC | Mitochondrial complex IV deficiency, nuclear type 5, (French-Canadian) |
| LTBP4 | Cutis laxa, autosomal recessive, type 1C |
| MAN2B1 | Alpha-mannosidosis |
| MCCC2 | 3-Methylcrotonyl-CoA carboxylase deficiency, type 2 |
| MCOLN1 | Mucolipidosis type 4 |
| MCPH1 | Microcephaly type 1 primary, autosomal recessive |
| MFN2 | Charcot-Marie-Tooth disease, axonal, type 2A2B |
| MFSD8 | Ceroid lipofuscinosis, neuronal, type 7 |
| MID1 | Opitz GBBB syndrome, type 1 |
| MKS1 | Bardet-Biedl syndrome type 13; Meckel syndrome, type 1; Joubert syndrome, type 28 |
| MLC1 | Megalencephalic leukoencephalopathy with subcortical cysts |
| MMAB | Methylmalonic aciduria, vitamin B12-responsive, type cblB |
| MMACHC | Methylmalonic aciduria and homocystinuria, cblC type |
| MMADHC | Homocystinuria, cblD type, variant 1 |
| MMUT | Methylmalonic aciduria, mut(0) type |
| MPDU1 | Congenital disorder of glycosylation, type 1F |
| MPI | Congenital disorder of glycosylation, type 1B |
| MPV17 | Mitochondrial DNA depletion syndrome type 6 (hepatocerebral); Charcot-Marie-Tooth disease, axonal, type 2EE |
| MPZ | Dejerine-Sottas disease |
| MTHFR | Homocystinuria due to MTHFR deficiency |
| MTM1 | Myotubular myopathy, X-linked |
| MTO1 | Combined oxidative phosphorylation deficiency 10 |
| MTR | Homocystinuria-megaloblastic anemia, cblG complementation type |
| MUSK | Fetal akinesia deformation sequence, type 1; Myasthenic syndrome, congenital, type 9 associated with acetylcholine receptor deficiency |
| MVK | Mevalonic aciduria |
| MYO15A | Deafness, autosomal recessive, type 3 |
| MYO5A | Griscelli syndrome, type 1 |
| MYO5B | Microvillus inclusion disease |
| MYO7A | Usher syndrome, type 1B; Deafness, autosomal recessive, type 2 |
| NAGA | Schindler disease, type I |
| NBN | Nijmegen breakage syndrome |
| NDP | Norrie disease |
| NEB | Nemaline myopathy type 2 |
| NEUROG3 | Diarrhea 4 malabsorptive, congenital |
| NHLRC1 | Epilepsy, progressive myoclonic, type 2B (Lafora) |
| NPC1 | Niemann-Pick disease, type C1 |
| NPC2 | Niemann-pick disease, type C2 |
| NPHP1 | Joubert syndrome type 4 |
| NPHP3 | Meckel syndrome type 7 |
| NPHP4 | Nephronophthisis type 4 |
| NPHS1 | Nephrotic syndrome, type 1 |
| NPHS2 | Nephrotic syndrome, type 2 |
| NR0B1 | Adrenal hypoplasia, congenital |
| NTRK1 | Insensitivity to pain, congenital, with anhidrosis |
| NUP62 | Striatonigral degeneration, infantile |
| OBSL1 | 3M syndrome 2 |
| OCA2 | Oculocutaneous albinism type 2 |
| OCRL | Lowe Syndrome; Dent disease type 2 |
| OPHN1 | Intellectual developmental disorder, X-linked syndromic, Billuart type |
| OTC | Ornithine transcarbamylase deficiency |
| PAH | Phenylketonuria |
| PAK3 | Intellectual developmental disorder, X-linked 30 |
| PC | Pyruvate carboxylase deficiency |
| PCCA | Propionic acidemia |
| PCDH15 | Deafness, autosomal recessive, type 23; Usher syndrome, type 1D/F digenic |
| PDHA1 | Pyruvate dehydrogenase E1-alpha deficiency |
| PDHB | Pyruvate dehydrogenase E1-beta deficiency |
| PDHX | Lacticacidemia due to PDX1 deficiency |
| PEX1 | Heimler syndrome type 1 |
| PEX10 | Peroxisome biogenesis disorder, type 6A (Zellweger syndrome); Peroxisome biogenesis disorder, type 6B |
| PEX16 | Peroxisome biogenesis disorder, type 8A (Zellweger syndrome); Peroxisome biogenesis disorder, type 8B |
| PEX7 | Rhizomelic chondrodysplasia punctata, type 1 |
| PGK1 | Phosphoglycerate kinase 1 deficiency |
| PHF8 | Intellectual developmental disorder, X-linked syndromic, Siderius type |
| PKHD1 | Polycystic kidney disease type 4 |
| PKP1 | Ectodermal dysplasia/skin fragility syndrome |
| PLA2G6 | Infantile neuroaxonal dystrophy type 1 |
| PLEC | Epidermolysis bullosa simplex with muscular dystrophy |
| PLOD1 | Ehlers-Danlos syndrome, kyphoscoliotic type, 1 |
| PLP1 | Pelizaeus-Merzbacher disease |
| PMM2 | Congenital disorder of glycosylation, type 1A |
| PNPLA1 | Ichthyosis, congenital, autosomal recessive, type 10 |
| PNPO | Pyridoxamine 5'-phosphate oxidase deficiency |
| POLG | POLG-related disorders |
| POLR1C | Leukodystrophy, hypomyelinating, type 11; Treacher Collins syndrome 3 |
| POMGNT1 | Muscular dystrophy-dystroglycanopathy, type 3A (Walker-Warburg syndrome); Type 3B; Type 3C (limb-girdle muscular dystrophy, type 15 [LGMDR15]) |
| POMT1 | Muscular dystrophy-dystroglycanopathy, type 1A (Walker-Warburg syndrome); Type 1B; Type 1C (limb-girdle muscular dystrophy, type 11 [LGMD R11]) muscular dystrophy, type 11 [LGMD R11]) |
| POMT2 | Muscular dystrophy-dystroglycanopathy, type 2A (Walker-Warburg syndrome); Type 2B; Type 2C (limb-girdle muscular dystrophy, type 14 [LGMD R14]) muscular dystrophy, type 14 [LGMD R14]) |
| POU1F1 | Pituitary hormone deficiency, combined, type 1 |
| POU3F4 | Deafness, X-linked, type 2 |
| PPT1 | Ceroid lipofuscinosis, neuronal, type 1 |
| PQBP1 | Renpenning syndrome |
| PRF1 | Hemophagocytic lymphohistiocytosis, familial, type 2 |
| PRICKLE1 | Epilepsy, progressive myoclonic, type 1B |
| PROP1 | Pituitary hormone deficiency, combined, type 2 |
| PRPS1 | Phosphoribosylpyrophosphate synthetase (PRS) deficiency |
| PSAP | Combined SAP deficiency |
| PYGL | Glycogen storage disease, type 6 |
| RAB3GAP1 | Warburg micro syndrome; Martsolf syndrome |
| RAB3GAP2 | Warburg micro syndrome; Martsolf syndrome |
| RAD51C | Fanconi anemia, complementation group O |
| RAG1 | Omenn syndrome; Severe combined immunodeficiency, B cell-negative |
| RAG2 | Omenn syndrome; Severe combined immunodeficiency, B cell-negative |
| RAPSN | Fetal akinesia deformation sequence, type 2; Myasthenic syndrome, congenital, type 11 associated with AChR deficiency |
| RARS2 | Pontocerebellar hypoplasia, type 6 |
| RELN | Lissencephaly 2 (Norman-Roberts type) |
| RMRP | Anauxetic dysplasia 1 |
| RNASEH2B | Aicardi-Goutieres syndrome, type 2 |
| ROR2 | Robinow syndrome, autosomal recessive |
| RP2 | Retinitis pigmentosa, type 2 X-linked |
| RPGR | Retinitis pigmentosa, type 3 X-linked; Cone-rod dystrophy, X-linked, 1 |
| RPGRIP1 | Leber congenital amaurosis, type 6 |
| RPGRIP1L | Joubert syndrome, type 7; Meckel syndrome, type 5; COACH syndrome |
| RS1 | Retinoschisis |
| RTEL1 | Dyskeratosis congenita, autosomal recessive type 5 |
| RYR1 | RYR1 related congenitalmyopathy |
| SACS | Spastic ataxia, Charlevoix-Saguenay, type |
| SBDS | Shwachman-Diamond syndrome |
| SCN4A | Myasthenic syndrome, congenital, type 16 |
| SCN9A | Indifference to pain and autosomal recessive hereditary sensory neuropathy type 2D |
| SCNN1B | Pseudohypoaldosteronism, type 1 |
| SCO2 | Mitochondrial complex IV deficiency, nuclear type 2 |
| SGCA | Limb-girdle muscular dystrophy, type 3 (LGMD R3) |
| SGCB | Limb-girdle muscular dystrophy, type 4 (LGMD R4) |
| SH3TC2 | Charcot-Marie-Tooth disease, type 4C |
| SIL1 | Marinesco-Sjogren syndrome |
| SLC12A1 | Bartter syndrome, type 1 |
| SLC12A6 | Agenesis of the corpus callosum with peripheral neuropathy |
| SLC16A2 | Allan-Herndon-Dudley syndrome |
| SLC17A5 | Salla disease |
| SLC19A2 | Thiamine-responsive megaloblastic anemia syndrome |
| SLC19A3 | Thiamine metabolism dysfunction syndrome, type 2 (biotin-or thiamine-responsive encephalopathy type) |
| SLC25A19 | Microcephaly, Amish type; Thiamine metabolism dysfunction syndrome 4 (progressive polyneuropathy type) |
| SLC25A22 | Developmental and epileptic encephalopathy 3 |
| SLC26A2 | Achondrogenesis, type 1B (diastrophic dysplasia) |
| SLC26A4 | Deafness, autosomal recessive, type 4; Pendred syndrome |
| SLC2A2 | Fanconi-Bickel syndrome |
| SLC35A1 | Congenital disorder of glycosylation, type 2F |
| SLC35C1 | Congenital disorder of glycosylation, type 2C |
| SLC37A4 | Glycogen storage disease, type 1B |
| SLC3A1 | Cystinuria |
| SLC45A2 | Albinism, oculocutaneous, type 4 |
| SLC4A11 | Corneal endothelial dystrophy, autosomal recessive |
| SLC6A8 | Cerebral creatine deficiency syndrome, type 1 |
| SMN1 | Spinal muscular atrophy |
| SMPD1 | Niemann-Pick disease, type A; Niemann-Pick disease, type B |
| SNAP29 | Cerebral dysgenesis, neuropathy, ichthyosis, and palmoplantar keratoderma syndrome |
| SP110 | Hepatic venoocclusive disease with immunodeficiency |
| SPATA7 | Leber congenital amaurosis, type 3 |
| SRD5A2 | 46,XY disorder of sex development due to 5-alpha-reductase 2 deficiency (pseudovaginal perineoscrotal hypospadias) |
| SRD5A3 | Congenital disorder of glycosylation, type 1Q; Kahrizi syndrome |
| ST3GAL5 | Salt and pepper developmental regression syndrome |
| STAR | Lipoid adrenal hyperplasia |
| STIL | Microcephaly, type 7 primary, autosomal recessive |
| STRA6 | Microphthalmia, isolated, with coloboma, type 8 |
| SUOX | Sulfite oxidase deficiency |
| SURF1 | Charcot-Marie-Tooth disease, type 4K; Leigh syndrome, due to COX IV deficiency |
| SYN1 | Epilepsy, X-linked, with variable learning disabilities and behavior disorders |
| TCTN1 | Joubert syndrome, type 13 |
| TCTN2 | Joubert syndrome, type 24; ?Meckel syndrome, type 8 |
| TF | Atransferrinemia |
| TGM1 | Ichthyosis, congenital, autosomal recessive, type 1 |
| TH | Segawa syndrome, recessive |
| THOC2 | intellectual developmental disorder, X-linked 12 |
| TJP2 | Cholestasis, progressive familial intrahepatic, type 4 |
| TMEM138 | Joubert syndrome 16 |
| TMEM216 | Joubert syndrome, type 2; Meckel syndrome, type 2 |
| TMEM237 | Joubert syndrome, type 14 |
| TMEM67 | Joubert syndrome, type 6; Meckel syndrome, type 3; COACH syndrome |
| TNNT1 | Nemaline myopathy, type 5 Amish type |
| TNXB | Ehlers-Danlos syndrome, classic-like |
| TPK1 | Episodic encephalopathy due to thiamine pyrophosphokinase deficiency |
| TPP1 | Ceroid lipofuscinosis, neuronal, type 2; Spinocerebellar ataxia, autosomal recessive, type 7 |
| TREX1 | Aicardi-Goutieres syndrome, type 1 |
| TRIM32 | Limb-girdle muscular dystrophy, type 8 (LGMD R8) |
| TSEN2 | Pontocerebellar hypoplasia, type 2B |
| TSEN34 | Pontocerebellar hypoplasia type 2C |
| TSEN54 | Pontocerebellar hypoplasia, type 2A; Pontocerebellar hypoplasia, type 4 |
| TSFM | Combined oxidative phosphorylation deficiency, type 3 |
| TTC21B | Short-rib thoracic dysplasia, type 4 with or without polydactyly |
| TTC8 | Bardet-Biedl syndrome, type 8 |
| TTN | Limb-girdle muscular dystrophy type 10 (LGMDR10); Early-onset myopathy with fatal cardiomyopathy (Salih myopathy) |
| TTPA | Ataxia with isolated vitamin E deficiency |
| TYR | Oculocutaneous albinism (OCA) type 1A; OCA type 1B |
| UBR1 | Johanson-Blizzard syndrome |
| UNC13D | Hemophagocytic lymphohistiocytosis, familial, type 3 |
| UPF3B | Intellectual developmental disorder, X-linked syndromic 14 |
| USH1C | Usher syndrome, type 1C; Deafness, autosomal recessive, type 18A |
| USH1G | Usher syndrome, type 1G |
| USH2A | Usher syndrome, type 2A |
| VLDLR | Cerebellar hypoplasia, impaired intellectual development, and dysequilibrium syndrome 1 |
| VPS13B | Cohen syndrome |
| VPS33B | Arthrogryposis, renal dysfunction and cholestasis, type 1 |
| WAS | Wiskott-Aldrich syndrome; Thrombocytopenia, X-linked |
| WDR62 | Microcephaly, type 2 primary, autosomal recessive, with or without cortical malformations |
| WFS1 | Wolfram syndrome, type 1 |
| WNT7A | Fuhrmann syndrome |
| WRN | Werner syndrome |
| XPA | Xeroderma pigmentosum, group A |
| XPC | Xeroderma pigmentosum, group C |
| ZAP70 | Autoimmune disease, multisystem, infantile-onset, type 2; Immunodeficiency, type 48 |
| ZDHHC9 | Intellectual developmental disorder, X-linked syndromic, Raymond type |
| ZMPSTE24 | Mandibuloacral dysplasia with, type B lipodystrophy |
| ZNF711 | Intellectual developmental disorder, X-linked 97 |

Мы создаём счастливые семьи
БЕЛГРАД
Karađorđeva 89
+381 11 414 65 65
info@cord.rs
НОВИ-САД
Fruškogorska 24
+381 21 31 01 333
info@cord.rs